Good Manufacturing Practice, or GMP, is the universal standard for quality production. It is a set of rules to ensure that products like medicines, food, and medical devices are made consistently and safely, batch after batch. The central idea is simple: quality cannot be inspected into a product at the end of the line. Instead, it ~ 해야 하다 be built into every step of the manufacturing process, from the raw materials that arrive at the loading dock to the final package that leaves it.
cGMP에서 'c'는 'Current(최신)'를 의미합니다. 이 한 글자는 중요하고도 역동적인 요구 사항을 제시합니다. GMP가 기본 지침을 제공한다면, cGMP는 제조업체가 현재 이용 가능한 최신 기술, 시스템 및 과학적 이해를 활용하도록 법적으로 의무화합니다. 10년 전 GMP 기준에 완벽하게 부합했던 공정이라도 그 이후로 더 나은, 더 신뢰할 수 있는 방법이 등장했다면 현재의 cGMP 검사에서는 불합격될 수 있습니다. 이는 기업들이 지속적으로 개선하도록 강제합니다.
핵심 요약
- GMP & cGMP distinction is now academic; the expectation is universal.
- Quality Risk Management (QRM) is the engine, not the paperwork nor the PLM.
- Data integrity is a primary audit focus.
- “Human Error” is a symptom, not a root cause. Attributing a deviation to “human error” is a red flag for a weak quality system.
- Process Analytical Technology (PAT) embodies the shift from testing to real-time assurance. The “c” in cGMP is exemplified by PAT.
- Supplier oversight is data-driven, not just audit-driven.
- The Qualified Person (QP) represents a critical EU-specific responsibility.
- 그만큼 Contamination Control Strategy (CCS) is the new cornerstone of sterile manufacturing.
The 10 Core Principles of Good Manufacturing Practice (GMP)
GMP is not just a set of rules but a quality mindset built upon ten fundamental principles. These principles work together to create a robust system that ensures quality is built into a product at every stage, rather than merely being tested for at the end.
1. Write Step-by-Step Procedures and Work Instructions
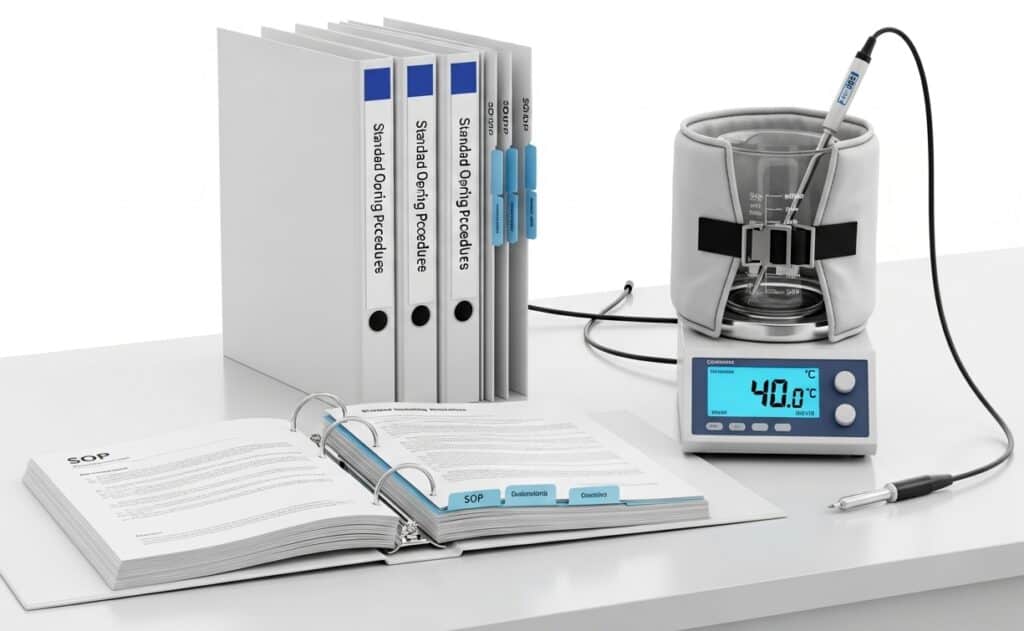
The foundation of GMP is ensuring that all processes are clearly defined and documented. This principle requires creating detailed, unambiguous Standard Operating Procedures (SOPs) for every critical task. The goal is to ensure that operations are performed consistently and correctly every time, regardless of who is performing the task. This eliminates ambiguity and provides a clear reference for training and execution.
Example of application: a company, “PharmaBlend Inc.,” manufactures a temperature-sensitive liquid drug. Their SOP for “Compounding Tank Temperature Control” (SOP-MFG-101) specifies not just the target temperature (40°C ± 2°C), but also the exact sequence for starting the heating jacket, the rate of temperature increase (not to exceed 5°C per minute), the specific calibrated probe to use for monitoring, and the actions to take if the temperature overshoots.
팁: instead of writing monolithic SOPs, use a modular approach. Create “master” SOPs for complex processes that reference smaller, task-specific “work instruction” documents for individual steps (e.g., calibrating a specific sensor, operating a single valve). This allows for easier updates—if a single piece of equipment is replaced, you only need to revise one small work instruction instead of the entire process SOP, significantly reducing review and approval time and minimizing the risk of introducing errors into unrelated sections.
2. Follow Procedures and Instructions Meticulously
Having documented procedures is meaningless if they are not followed. This principle demands strict adherence to the written SOPs without deviation. If a deviation is necessary, it must be formally documented, justified, and approved through a defined change control process. This ensures that any departure from the standard is controlled, assessed for risk, and recorded for traceability.
팁: implement a “Right-First-Time” (RFT) metric for procedure execution, tracked during batch record review. When deviations occur due to non-adherence, don’t just retrain the operator. Perform a root cause analysis focused on the procedure’s 사용성 (a Human Factors approach). Was the instruction ambiguous? Was the sequence illogical? Is the required tool hard to access? Improving the procedure itself is a more effective long-term Corrective and Preventive Action (CAPA) than simply blaming human error.
3. Promptly and Accurately Document Work
This is the principle of “if it wasn’t written down, it didn’t happen.” All activities, from receiving raw materials to shipping the final product, must be documented in real-time. This includes recording data, signatures, dates, and any observations. Accurate, contemporaneous documentation provides a complete and traceable history of a batch (known as a Batch Record or Device History Record), which is essential for investigating deviations, troubleshooting problems, and proving compliance during an audit.
팁: when designing batch records (paper or electronic), incorporate “data integrity checks” directly into the fields. For example, instead of just a blank space for “End Time,” structure it to require a start time and an end time, with an automated or manual check to ensure the duration is logical for the process step. For critical entries, use “verify-by-second-person” sign-offs, but ensure the verifier is trained to re-perform the critical calculation or check the setting, not just “check the box.”
Difference between DMR and DHR:
- The Device Master Record, or DMR, is the master recipe for a medical device.
- 이는 일관된 제품을 생산하는 데 필요한 모든 지침, 사양 및 절차를 공식적으로 통제하여 편집한 것입니다. DMR에는 설계 도면 및 재료 사양부터 자세한 제조 지침, 품질 관리 테스트 방법, 라벨링 및 포장 요구 사항에 이르기까지 모든 것이 포함되어 있습니다.
완전한 청사진이라고 생각하세요.
Validation is the documented proof that a process, system, or piece of equipment consistently produces the expected result. This principle requires manufacturers to prove that their processes are reliable and under control. This includes validating manufacturing processes, analytical testing methods, cleaning procedures, and computerized systems to ensure they are fit for their intended purpose.
실제로는, IQ OQ PQ process validation is how a company produces that documented proof: installation, operational and performance checks of each system are signed off before routine production starts. Smaller sites often begin with IQ and OQ validation of new equipment, then complete the performance step during the first production batches.
팁: adopt a lifecycle approach to validation based on ASTM E2500. Instead of treating validation as a one-time event, integrate it with Quality by Design (QbD). Define a “control space” during process development, and use the validation exercise (Process Performance Qualification – PPQ) not just to confirm the process works, but to verify that it remains within this state of control. This shifts the focus from a simple pass/fail event to demonstrating ongoing process understanding and control, which is highly valued by regulators.
For this stage, we suggest our very complete reading on IQ OQ PQ 공정 검증:
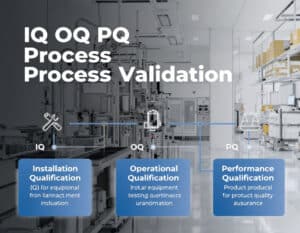
Expert tip: validation is becoming a continuous lifecycle, not a one-time event. The “three successful batches” mindset is becoming obsolete. cGMP demands a lifecycle approach to ensure the process remains in a state of control for its entire commercial life:
- a robust process design (Stage 1)
- formal qualification (Stage 2)
- 그리고 지속적인 공정 검증 프로그램(3단계)
5. Properly Design, Build, and Maintain Facilities and Equipment
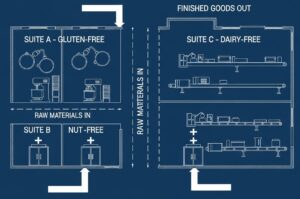
The physical environment and the tools used are critical to product quality. This principle dictates that facilities must be designed to prevent cross-contamination and mix-ups. Equipment must be designed for its intended use, easy to clean, and properly calibrated and maintained.
The layout, airflow, and material flow should all be logically designed to protect the product such as unidirectional flow for personnel and materials. Raw materials enter one end, move through dedicated processing suites with positive air 압력 differentials, and finished goods exit the other end. There are no crossing paths between raw and finished goods, and personnel must pass through airlocks to enter production areas, minimizing the risk of cross-contamination.
팁: when designing a new facility or modifying an old one, use 3D modeling and virtual reality (VR) walkthroughs with a cross-functional team (including operators, maintenance, and QA) before construction begins. This allows you to identify ergonomic issues, inefficient material flows, and hard-to-clean areas (e.g., inaccessible pipework, awkward equipment placement) at the design stage, where changes are cheap, rather than after the facility is built, where they are prohibitively expensive.
6. Maintain Good Housekeeping and Hygiene
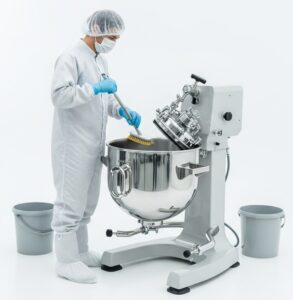
Contamination is a major risk in manufacturing. This principle requires maintaining a high level of cleanliness and hygiene throughout the facility. This includes personal hygiene standards for employees (e.g., proper gowning), documented cleaning and sanitation schedules for facilities and equipment, and pest control programs. The goal is to protect the product from physical, chemical, and microbial contaminants.
Example of application: the cleaning procedure for a mixing vessel is not just “wash with detergent.” It’s a validated, multi-step process: pre-rinse with purified water, wash with a specific concentration of a validated cleaning agent for a set time, final rinse with Water-For-Injection (WFI), and finally, a swab test for chemical and microbial residues to prove the vessel is truly clean before the next batch can be made.
참고: 청소는 소독과 다릅니다. (refer to other articles on this).
팁: implement a robust environmental monitoring (EM) program that goes beyond routine sampling. Use EM data to create trend analysis charts and heat maps of your facility. A spike in microbial counts in one area isn’t just a deviation to be closed; it’s a data point. Use this data proactively to identify “hot spots,” assess the effectiveness of cleaning procedures over time, and adjust sanitation frequencies based on risk and data, rather than a fixed, arbitrary schedule.
7. Build Quality into the Entire Product Lifecycle
This principle emphasizes that quality is not just the responsibility of the Quality Control (QC) department. It must be integrated into every stage, from research and development, through raw material sourcing (supplier qualification), manufacturing, packaging, and distribution. It involves a holistic approach where every department understands its role in maintaining product quality.
Example of application: “Precision Pills LLC” is developing a new tablet. During the R&D phase (long before manufacturing), they use Quality by Design (QbD) principles. They identify Critical Quality Attributes (CQAs) like tablet hardness and dissolution rate. They then perform experiments to understand how Critical Process Parameters (CPPs) like turret speed and compression force affect these CQAs. This knowledge is then transferred to manufacturing, ensuring a robust and well-understood process from day one.
팁: establish a formal Technology Transfer team that acts as a bridge between R&D, Engineering, and Manufacturing. This team should manage a “knowledge transfer package” that includes not just the process parameters, but the rationale behind them (the “why”). This package should detail failed experiments, process boundaries, and the scientific understanding of the product. This prevents manufacturing from treating the process like a “black box” and enables more effective troubleshooting and continuous improvement later on.
8. Perform Quality Control and Audits
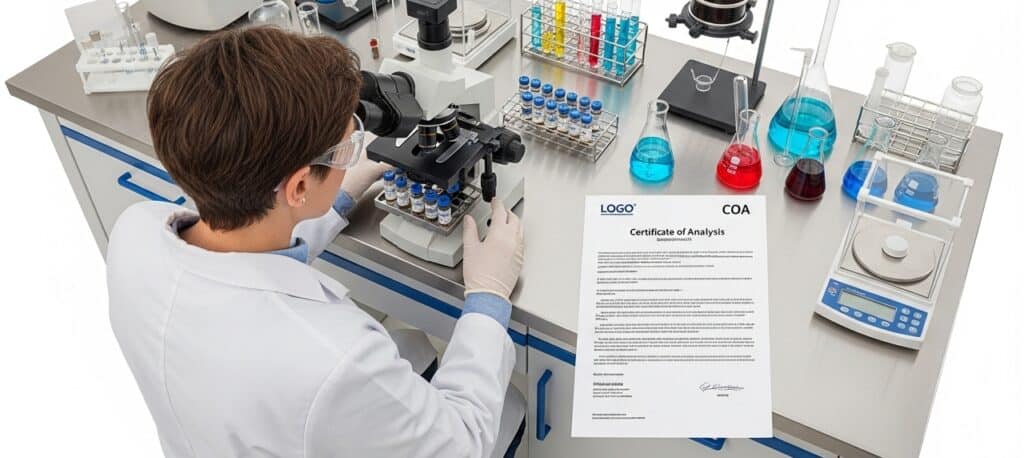
An independent Quality unit (comprising Quality Assurance and Quality Control) is essential. This principle ensures that proper controls are in place. This includes testing raw materials, in-process samples, and finished products against pre-defined specifications. It also involves conducting regular internal audits (self-inspections) to assess the effectiveness of the GMP system and ensure it is being followed correctly.
Example of application: “SafeInjectables Corp.” receives a shipment of vials from a new supplier. Even though the supplier provided a Certificate of Analysis (CoA) showing the vials meet all specifications, the internal QC lab performs its own independent identity and critical defect testing (e.g., for cracks and dimensions) on a sample of the vials before the shipment is released for use in production. This verifies the supplier’s data and protects against potential quality issues.
팁: structure your internal audit program to be process-based, not just department-based. Instead of auditing the “Warehouse Department,” conduct an audit of the “Material Control Process” which would follow the material from receiving, through warehouse storage, dispensing, and return-to-stock. This approach breaks down departmental silos and provides a much clearer picture of the health and efficiency of the 엔드투엔드 process, revealing risks that occur at the hand-off points between departments.
9. Protect Products Against Contamination
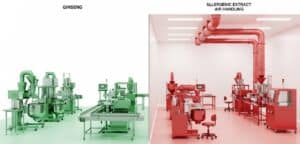
This is a central theme that overlaps with other principles but deserves its own focus. It involves designing processes and facilities to prevent contamination of the product with any foreign substance.
This includes preventing cross-contamination between different products, microbial contamination, and contamination from personnel or the environment.
Measures like closed production systems, proper gowning, and controlled material handling are key applications of this principle.
Example of application: a facility, “MultiHerb Supplements,” produces both a ginseng supplement and a potent herbal extract known to be an allergen. To prevent cross-contamination, the allergenic extract is produced in a completely segregated suite with its own dedicated air handling system (HVAC) and equipment. All tools are color-coded red and are never allowed to leave the suite. This physical and procedural separation is critical to protecting other products.
팁: 물리적 분리를 넘어 독성학적 데이터를 기반으로 한 공용 장비의 '세척 검증' 전략을 시행하십시오. 각 제품에 대해 일일 허용 노출량(PDE) 값을 계산하십시오. 이 PDE 값을 사용하여 제품 잔류물에 대한 과학적으로 타당하고 건강 기반의 세척 한계를 설정하십시오. 이러한 위험 기반 접근 방식은 현재 업계 표준(EMA에서 권장)이며 '10ppm' 또는 '육안으로 깨끗함'과 같은 기존의 임의적인 한계에 의존하는 것보다 훨씬 더 효과적입니다.
10. Train and Develop Competent Personnel
The human element is often the most critical and variable factor in manufacturing. This principle requires that all employees are properly trained for their specific roles. Training should cover not only the technical aspects of their 직업 (how to operate equipment) but also the principles of GMP and the potential consequences of not following procedures. Competency must be regularly assessed and documented.
Example of application: before an operator at “CellTherapy Innovations” is allowed to work independently on the critical cell culture expansion step, they must complete a multi-stage qualification program. This includes reading SOPs, observing a qualified operator, performing the task under direct supervision, and finally, successfully processing several “test” batches on their own. Their competency is documented and certified by their supervisor and the QA department in their official training file ( mandatory in most of the sectors where GMP applies).
팁: 단순한 '교육 기반' 시스템에서 '역량 기반' 시스템으로 전환하십시오. 작업자가 표준 작업 절차(SOP)에 대한 '교육'을 받았다는 사실만 문서화하는 대신, 작업자의 기술과 지식을 입증하도록 요구하는 공식적인 역량 평가를 개발하십시오. 여기에는 실기 시험(예: '정확하게 조립하고')이 포함될 수 있습니다. 분해하기 이 주입 펌프 작동법)과 구두 평가("이 단계의 핵심 매개변수를 설명하고, 매개변수가 벗어날 경우 어떻게 할 것인지 설명해 주세요.")로 구성됩니다. 이를 통해 자격 검증 기록이 훨씬 더 견고해지고, 단순히 출석하는 것이 아니라 진정한 이해를 보장할 수 있습니다.
GMP의 범위와 응용
The scope of GMP is broad, covering any industry where the quality of the product can have a significant impact on consumer health and safety. The application of GMP principles is tailored to the specific risks associated with each industry.
1. Pharmaceuticals and biotechnology 처방약, 일반의약품을 포함한 모든 의약품 제조를 포괄합니다. 백신및 활성 의약품 성분(API). This is the most stringent application of GMP. It involves strict control over raw materials, aseptic (sterile) processing for injectables, rigorous process validation, stability testing to determine shelf life, and a “Qualified Person” in the EU responsible for certifying that each batch meets all legal and quality requirements before release. 이 분야는 21 CFR Part 211(미국) 및 EudraLex Volume 4(유럽)와 같은 규정의 적용을 받습니다. | 2. Medical devices Includes everything from simple tongue depressors to complex life-sustaining equipment like pacemakers and MRI machines. The focus is on design controls, ensuring the device is designed to be safe and effective from the outset. GMP for medical devices (often called the Quality System Regulation or QSR) emphasizes risk management (ISO 14971), traceability of components, and maintaining a complete Device Master Record (DMR) and Device History Record (DHR). The key regulation in the US is 21 CFR Part 820. |
3. Food and beverage Covers the processing, packaging, and holding of human food. GMP in the food industry focuses on preventing contamination and ensuring food safety. Key applications include 위해 요소 분석 및 중요 관리점 (HACCP) systems to identify and control food safety hazards, allergen management programs to prevent cross-contact, sanitation procedures, and pest control. The Food Safety Modernization Act (FSMA) in the US heavily incorporates cGMP principles. |  |
4. Cosmetics Includes products like makeup, lotions, shampoos, and soaps. While often less stringent than for pharmaceuticals, GMP for cosmetics focuses on preventing microbial contamination, ensuring product stability, and accurately labeling ingredients. The ISO 22716 standard provides specific GMP guidelines for the cosmetic industry, covering production, control, storage, and shipment. | 5. Dietary supplements Covers vitamins, minerals, herbs, and other supplements. GMP ensures that supplements are produced without contaminants, are accurately labeled, and contain the ingredients they claim to. This involves identity testing of raw materials, ensuring proper formulation, and controlling for contaminants like heavy metals and pesticides. In the US, this is governed by 21 CFR Part 111. |
The Qualified Person (QP)
In the European Union, the Qualified Person (QP) serves as the final, legally mandated gatekeeper for every batch of medicinal product.
Before a batch can be released for sale or for use in a clinical trial, a QP must personally certify that it complies with all regulatory requirements. This certification is a formal attestation that the batch was manufactured and tested in accordance with its specific 마케팅 Authorisation and the principles of Good Manufacturing Practice (GMP). This is not a corporate sign-off; it is a profound personal and legal responsibility placed upon a named individual, who must be a registered professional, such as a pharmacist or chemist, with extensive practical experience.
The QP’s responsibilities extend far beyond the final review of a batch record. They must ensure that the entire Pharmaceutical Quality System is functioning correctly. This includes verifying that all starting materials are from qualified supply chains, that manufacturing and testing processes are properly validated, that all deviations and changes have been appropriately investigated and approved, and that all necessary audits have been performed.
The QP does not personally perform every task, but they are ultimately accountable for the systems that do. They must have a comprehensive understanding of the entire manufacturing and control process, with the authority to access any relevant area or document and the power to halt a release if any aspect of compliance is in doubt.
cGMP에 특정한 추가 요구 사항
GMP의 기본 원칙을 바탕으로 구축된 cGMP는 더욱 역동적이고, 능동적이며, 기술적으로 진보된 일련의 요구 사항을 도입합니다.
'Current'에 대한 'c'는 단순한 제안이 아니라 제조업체의 시스템이 최첨단이어야 한다는 법적 및 규제적 기대 사항입니다.
1. 사전 예방적 품질 위험 관리(QRM)
While basic GMP implicitly involves managing risk, cGMP mandates the use of a formal, systematic, and proactive process for managing risks to product quality throughout the product lifecycle. This is not just about reacting to failures but about anticipating and preventing them. This principle is heavily guided by the international standard ICH Q9.
Example of application: 허구의 �8220;넥스트젠 바이오로직스이 회사는 새로운 단일클론 항체를 개발하고 있습니다. 공정 개발 과정에서 공식적인 고장 모드 및 영향 분석(FMEA)을 수행합니다.FMEA그들은 생물반응기 공정에서 '세균 오염'을 위험도가 높은 심각한 실패 모드로 식별했습니다. 이러한 위험을 사전에 제어하기 위해, 복잡하고 위험한 현장 증기 멸균(SIP) 검증이 필요한 기존의 스테인리스 스틸 반응기 대신 일회용 멸균 생물반응기 백을 사용하는 공정을 설계했습니다. 이러한 설계 선택은 QRM 공정의 직접적인 결과입니다(플라스틱 폐기물 제거는 여기서의 주제가 아닙니다).
팁: Integrate your QRM system directly into your Change Control and Deviation Management systems. For every change or deviation, require a documented risk assessment that determines the scope of validation, testing, and regulatory action needed. This creates a scientifically sound and legally defensible rationale for your decisions, allowing you to justify why a minor change only required limited testing, while a major change required a full re-validation and submission to a regulatory agency.
2. 의약품 품질 시스템(PQS) 구현
cGMP는 단순히 일련의 품질 관리 절차 이상의 것을 요구합니다. GMP 원칙과 회사의 사업 목표를 연계하여 지속적인 개선을 이끌어내는 포괄적이고 통합적인 관리 시스템을 필요로 합니다.
ICH의 지침에 따라 Q10PQS는 고위 경영진이 품질에 적극적으로 참여하고 책임을 지도록 보장합니다.
팁: PQS의 효과성을 감사자에게 입증하려면 품질 목표를 핵심 성과 지표(KPI)와 직접 연결하십시오. 단순히 '편차 건수'만 추적하는 대신 '편차 재발률'을 추적하십시오. 낮은 재발률은 강력한 증거입니다. KPI 이는 귀사의 근본 원인 분석 및 CAPA 시스템이 효과적임을 입증합니다. 예방하다 문제점을 해결하는 것이 바로 성숙한 PQS의 핵심 목표입니다.
3. 공정 분석 기술(PAT) 및 고급 공정 제어의 활용
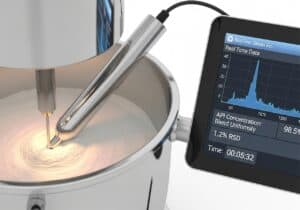
이것은 '커런트' 철학의 초석입니다.
cGMP는 제조업체가 완제품 샘플 테스트에만 의존하는 방식에서 벗어나도록 권장하고 있으며, 점점 더 그러한 방식을 요구하고 있습니다. 제조업체는 최신 기술을 활용하여 제조 공정을 실시간으로 모니터링하고 제어함으로써 모든 단계에서 제품의 품질을 확보해야 합니다.
Example of application: while a traditional GMP process for tablet blending involves mixing powders for a fixed time (e.g., 20 minutes) and then taking a few samples to a lab for testing, the cGMP approach would use an in-line Near-Infrared (NIR) 분광학 probe directly inside the blender. The probe continuously measures the blend’s uniformity in real-time. The blending process is stopped not based on time, but at the precise moment the NIR data confirms the blend has achieved the validated state of homogeneity, ensuring perfect results for every batch.
팁: 새로운 PAT 장비를 도입할 때는 처음 10~20개 배치에 대해 '섀도잉' 방식을 사용하는 것이 좋습니다. PAT 시스템을 가동하여 실시간 데이터를 수집하되, 모든 품질 관련 결정(예: 배치 출하 승인)은 기존의 검증된 오프라인 방식을 기반으로 계속 진행하십시오. 이를 통해 견고한 데이터 모델을 구축하고 위험 부담이 적은 환경에서 PAT 시스템의 신뢰성을 입증한 후, 변경 관리 절차를 통해 기존의 비효율적인 방식을 대체하는 공식적인 검증을 진행할 수 있습니다.
4. 타협 없는 데이터 무결성 및 21 CFR 파트 11 준수
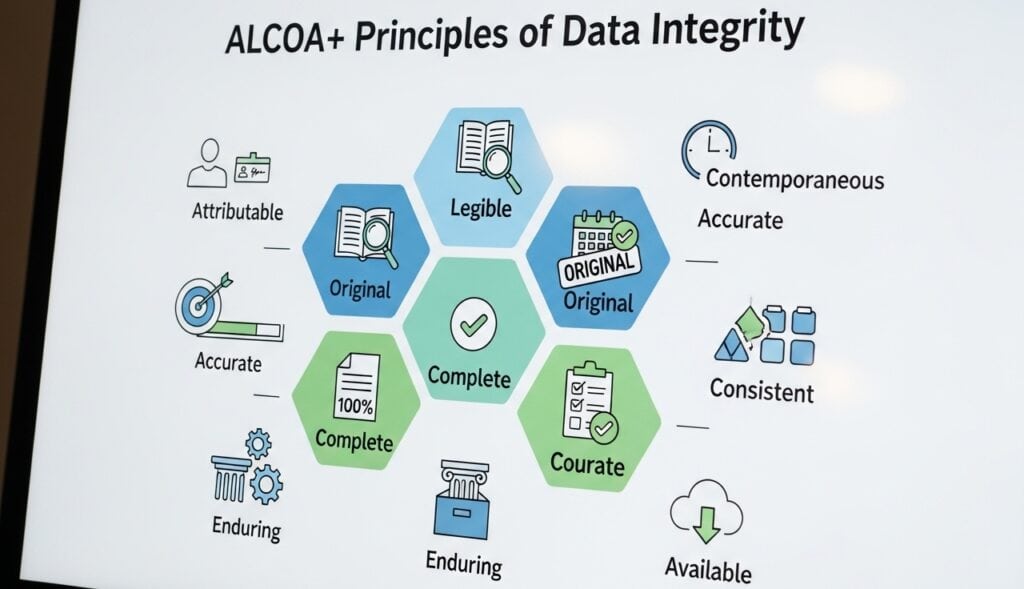
GMP는 항상 정확한 기록을 요구해 왔지만, cGMP는 기록의 무결성에 더욱 중점을 둡니다. 전자 데이터. 컴퓨터 시스템의 등장과 함께,
규제 기관은 데이터가 전체 수명 주기 동안 우발적 또는 의도적인 변경, 삭제 또는 손실로부터 완벽하게 보호되어야 한다는 절대적인 보장을 요구합니다. 이는 흔히 ALCOA+ 원칙(귀속성, 가독성, 동시성, 원본성, 정확성, 그리고 완전성, 일관성, 지속성, 가용성)으로 요약됩니다.
Example of application: 'DataSecure Labs'의 연구실에서는 고성능 액체 크로마토그래피(HPLC) 시스템을 사용합니다. 기본적인 GMP 설정에서는 독립형 PC에서 결과를 출력할 수 있습니다. cGMP를 준수하는 설정에서는 HPLC 소프트웨어가 네트워크에 연결되어 있으며 21 CFR Part 11을 완벽하게 준수합니다. 분석가는 고유한 사용자 이름과 비밀번호 없이는 로그인할 수 없습니다. 모든 원시 데이터는 보안 서버에 직접 저장되며 삭제하거나 덮어쓸 수 없습니다. 적분 매개변수 변경을 포함한 모든 작업은 수정할 수 없는 영구적인 타임스탬프 감사 추적 기록에 자동으로 기록됩니다.
팁: '부정적 테스트' 또는 '적대적 감사'를 실시하여 시스템에 대한 적극적인 검증을 진행하십시오. 기술에 정통한 팀원에게 시스템의 데이터 무결성을 의도적으로 손상시키도록 지시하십시오. 분석 날짜를 소급하여 시스템 시계를 변경할 수 있습니까? 감사 기록에 남지 않도록 실패 결과를 삭제할 수 있습니까? 로그인 시스템을 우회할 수 있습니까? 이러한 연습은 노련한 규제 기관이 조사할 법한 상황을 모방한 것으로, 취약점을 찾아 수정하는 가장 빠른 방법입니다.
5. 지속적인 개선을 보여주는 문화
cGMP는 본질적으로 고정된 목표가 아니라 지속적인 여정입니다. 규제 기관은 모든 출처(공정 모니터링, 일탈, 불만 사항, 내부 감사)의 데이터를 적극적으로 수집, 추세 분석하고 이를 활용하여 공정과 시스템을 의미 있게 개선하는 공식적인 시스템을 기대합니다.
목표는 시간이 지남에 따라 발전하고 더욱 효율적이고 신뢰할 수 있게 되는 것입니다.
cGMP 접근 방식의 예: 제조 라인의 밀봉 기계에서 반복적으로 경미한 경보가 발생합니다. 기본적인 GMP 대응은 경보를 기록하고 매번 재설정하는 것입니다. cGMP 대응은 경보가 세 번 발생한 후, 제품 품질 시스템(PQS)이 자동으로 공식적인 시정 및 예방 조치(CAPA)를 발동하는 것입니다. 여러 부서로 구성된 팀이 추세를 조사하고, 근본 원인이 공급업체의 호일 재질에 미세한 편차가 있음을 파악한 후, 공급업체와 협력하여 재질 사양을 강화합니다. 그런 다음, 해결책의 효과를 검증하기 위해 향후 6개월 동안 경보 발생률을 모니터링합니다.
팁: 연간 보고서를 기반으로 공식적인 '피더' 시스템을 구축하세요. 제품 리뷰 연간 품질 검토(APR) 또는 제품 품질 검토(PQR)를 지속적인 개선 프로그램에 직접 통합하십시오. PQR이 단순히 서류철에 놓이는 문서가 되지 않도록 하십시오. 모든 PQR에 다음 해에 적용할 수 있는 구체적이고 데이터 기반의 프로세스 개선 또는 위험 감소 기회를 최소 하나 이상 명시하도록 의무화하십시오. 이를 통해 규정 준수와 선제적이고 미래지향적인 프로세스 개선을 공식적으로 연계할 수 있습니다.
GMP와 cGMP 비교
| GMP | cGMP | |
| 핵심 원칙 및 목표 | A comprehensive quality system to ensure products are consistently produced and controlled according to established standards. The fundamental goal is to build quality into the 제품 디자인 and manufacturing process to minimize risks that cannot be eliminated by testing the final product alone. | |
| 핵심 강조 사항 | 품질 제조를 위한 기본적이고 국제적으로 인정받는 최소 요구 사항을 확립합니다. 이는 기준 표준입니다. | 'C'는 제조업체가 최신 기술, 시스템 및 과학적 접근 방식을 사용하도록 법적으로 의무화하는 규정입니다. 이는 선제적으로 발전하는 표준입니다. |
| 표준의 본질 | 안정적이고 체계화된 기준선을 제공합니다. 주기적으로 업데이트되지만, 확립되고 문서화된 규칙 준수에 중점을 둡니다. | 역동적이고 끊임없이 진화하는 환경입니다. 기업은 업계 동향을 적극적으로 모니터링하고 최신 품질 보증 시스템을 도입하여 규정을 준수해야 합니다. 어제 규정을 준수했던 것이 내일은 그렇지 않을 수도 있습니다. |
| 주요 규정 및 관할 기관 | 이 용어는 전 세계적으로 사용되며, 통일된 국제 표준을 나타냅니다. | 이 특정 용어는 미국 식품의약국(FDA)에서 시행하는 공식적인 법적 기준입니다. 하지만 '최신 정보'를 제공하는 원칙은 대부분의 주요 규제 기관에서 요구하는 사항입니다. |
| 유럽 연합 | 유럽의약품청(EMA)은 EudraLex §4를 통해 GMP(우수 의약품 제조 및 품질관리 기준)를 시행하며, 이 지침서는 인체 및 동물용 의약품에 대한 상세한 가이드라인을 제공합니다. | 'cGMP'라는 용어는 사용되지 않지만, 최신 기준을 준수해야 한다는 기대는 EudraLex 내부에 내재되어 있으며 검사 과정에서 엄격하게 시행됩니다. |
| 미국 | 미국 식품의약국(FDA)은 cGMP라는 용어를 사용하여 규정이 유연하며 최신 기술을 요구한다는 점을 강조합니다. 주요 규정은 다음과 같습니다.
| |
| 국제기구 | 세계보건기구(WHO)는 전 세계적인 GMP(우수 의약품 제조 및 품질관리 기준)를 제정하며, 자체적인 광범위한 규제 체계가 없는 국가들이 이를 채택하는 경우가 많습니다. 국제 의약품 규제 조화 위원회(ICH)는 유럽 연합, 미국, 일본 간의 기준을 조화시키기 위한 가이드라인(예: 원료의약품에 대한 ICH Q7)을 개발합니다. | 이러한 단체들은 cGMP 철학의 핵심인 지속적인 개선과 현대적인 품질 시스템 원칙을 장려합니다. |
| 적용 분야 |
| |
| 기업에 미치는 실질적인 영향 | 체계적이고 문서화된 시스템 구축에 중점을 둡니다. 여기에는 표준 운영 절차(SOP), 직원 교육, 장비 검증 및 꼼꼼한 기록 관리가 포함됩니다. | GMP(우수 제조 기준)의 모든 요건을 충족하는 것은 물론, 지속적인 개선을 위한 적극적인 문화가 필요합니다. 여기에는 고급 공정 제어, 최신 분석 시험 방법, 강력한 품질 위험 관리(QRM), 그리고 포괄적인 시정 및 예방 조치(CAPA) 시스템 구현이 포함됩니다. |
| 비용 및 투자 | 여기에는 규정을 준수하는 품질 시스템, 시설 및 인력 교육을 구축하고 유지하는 데 필요한 표준 비용이 포함됩니다. | 최첨단 기술을 유지하기 위해서는 새로운 기술, 고급 분석 장비, 지속적인 검증 연구 및 전문 교육에 대한 더 높고 지속적인 투자가 필요한 경우가 많습니다. |
상세 제조 예시 (무균 충전) | 프로세스: 기존의 A/B 등급 클린룸에 있는 오래된 무균 충전 라인입니다. 작업자는 가운을 완전히 착용하고 빈번하게 수동 작업(예: 스토퍼 장착, 막힘 제거)을 수행합니다. 환경 모니터링은 배치 시작과 종료 시점에 침전판 검사 및 공기 샘플링을 통해 주기적으로 실시됩니다. 규제: 이전 GMP 규정의 서면 내용(예: EudraLex Annex 1의 이전 버전)을 준수합니다. 확인: 해당 프로세스는 수년 전에 성공적인 미디어 게재를 통해 검증되었으며, 재검증은 최신 위험 기반 접근 방식 없이 정해진 일정(예: 2년마다)에 따라 수행됩니다. 선적 서류 비치: 꼼꼼한 종이 기반 배치 기록에는 모든 단계, 개입 사항 및 주기적인 모니터링 결과가 문서화되어 있습니다. (수동 개입으로 인한 오염 위험이 높고 현재 기술로는 지속적인 실시간 공정 데이터 확보가 불가능하여 cGMP 기준을 충족하지 못할 수 있음) | 프로세스: 완전 자동화된 접근 제한 장벽 시스템(RABS) 또는 격리 장치를 사용하는 최첨단 무균 충전 라인입니다. 로봇 시스템이 모든 자재 이송을 처리하며, 모든 작업은 글러브 포트를 통해서만 이루어져 사람과의 접촉을 최소화합니다. 지속적인 실시간 입자 모니터링 시스템이 통합되어 환경 이상 발생 시 즉각적인 경보를 제공합니다. 규제: 이는 위험 기반 접근 방식과 첨단 기술 사용을 통해 무균성을 보장하도록 규정하는 현재 FDA의 기대치 및 개정된 EudraLex Annex 1과 일치합니다. 확인: 해당 공정은 오염 위험을 사전에 식별하고 완화하는 품질 위험 관리(QRM) 평가(ICH Q9)를 기반으로 검증되었습니다. 공정 분석 기술(PAT)을 사용하여 주요 매개변수를 실시간으로 모니터링할 수 있습니다. 선적 서류 비치: 규정을 준수하는 전자 배치 기록(21 CFR Part 11)은 모든 공정 매개변수와 지속적인 모니터링 데이터를 자동으로 캡처하여 지속적인 관리 상태를 입증하는 완전하고 실시간이며 감사 가능한 데이터 기록을 제공합니다. |
결론
실제로 GMP와 cGMP의 구분은 모호한데, 두 시스템 모두 제품의 안전성과 효능을 일관되게 보장한다는 동일한 기본 목표를 공유하기 때문입니다. GMP는 필수적인 원칙, 즉 영구적인 규칙을 제시합니다. cGMP의 'c'는 이러한 원칙을 최신 기술과 과학적 이해를 바탕으로 반드시 준수해야 한다는 필수적인 요건을 명확히 하는 것입니다.
Documents in this field are easier to read with a list of manufacturing terms and acronyms such as GMP, cGMP and PQS at hand.
이는 더 이상 미국 FDA만의 개념이 아닙니다. 전 세계 규제 기관들은 공식 문서에서 'GMP'만을 언급하더라도 이제는 보편적으로 이를 현행 우수 의약품 제조 기준(CMP)으로 해석하고 있으며, 감사관은 당시 기준을 얼마나 잘 준수했든 간에 시대에 뒤떨어진 프로세스는 받아들이지 않고, 자체 평가 및 실시간 개선이 가능한 프로세스를 요구합니다.
본질적으로 cGMP는 GMP를 현대 과학에 맞춰 발전시켜 제품 안전과 품질을 보장하는 것입니다.
자주 묻는 질문
In a practical audit, how does an inspector’s expectation for ‘cGMP’ differ from the written ‘GMP’ regulations?
An inspector expects to see not just that you follow your written procedures (GMP), but that your procedures themselves reflect current industry best practices and technology (cGMP). They will question why you are using a 20-year-old analytical method when a more accurate and reliable one is now standard, or why you rely on manual checks where automated in-line verification is now common. They are auditing your awareness and proactive adoption of modern quality standards.
Does cGMP mean we must constantly invest in the newest technology, or can we justify using older, validated equipment?
You can absolutely justify using older equipment, but the burden of proof is on you. Your justification must be documented and risk-based. You need to demonstrate through robust validation, rigorous maintenance, intensive monitoring, and trend data that your older system provides an equivalent or superior level of quality assurance and 프로세스 제어 compared to modern alternatives. If your process using old equipment has a higher deviation rate, you will not be able to defend it.
Beyond audit trails, what are the most common ‘unseen’ data integrity gaps regulators are focusing on?
Regulators are heavily scrutinizing uncontrolled spreadsheets used for GMP calculations, the use of shared login credentials on standalone equipment (like balances or pH meters), and the ability to perform “test runs” on analytical equipment that can be deleted without a trace. Another major focus is the integrity of metadata—the data about the data, such as timestamps and user IDs, which must be securely linked to the original record.
How is a Pharmaceutical Quality System (PQS) under ICH Q10 different from just having a strong QA department?
A strong QA department enforces quality; a PQS manages it as a business-wide objective. The key difference is the formal integration of senior management and a focus on process performance and continuous improvement. A PQS ensures that quality metrics directly influence business decisions (like resource allocation and strategic planning) and that management is actively reviewing and driving the system’s effectiveness, as opposed to delegating all quality matters to QA.
What does a “lifecycle approach” to process validation (per ASTM E2500) actually mean for an engineer?
It means validation is no longer a “three-and-done” batch exercise. It’s a continuous process. For an engineer, this means: Stage 1 (Process Design): using Quality by Design (QbD) to define a robust process and its control space. Stage 2 (Process Qualification): verifying the facility and equipment are fit for purpose and that the process consistently works within its defined space (PPQ). Stage 3 (Continued Process Verification): actively monitoring the process during routine production using statistical process control (SPC) to ensure it remains in a state of control for its entire commercial life.
What is the most significant practical difference between EU GMP (EudraLex) and US cGMP?
The most significant difference is the role of the Qualified Person (QP) in the EU. In the US, the Quality Unit has the authority to release a batch. In the EU, a specifically named QP must personally certify that each batch has been manufactured and tested in accordance with all regulations and the marketing authorization before it can be released. This places an immense personal and legal responsibility on one individual.
Our CAPA system is compliant, but problems often recur. What is the cGMP expectation for “CAPA effectiveness”?
The cGMP expectation is that you formally verify your CAPAs worked. This requires building an “effectiveness check” step into your CAPA procedure. This check, performed weeks or months after the CAPA is implemented, must provide objective data (e.g., trend analysis of deviation rates, new audit findings) to prove that the root cause was eliminated and the problem has not recurred. A CAPA closed without this verification is a major red flag for auditors.
How has the cGMP expectation for supplier qualification evolved beyond just auditing the supplier?
Audits are still necessary, but cGMP now expects a more data-driven, risk-based approach. This includes establishing formal Quality Agreements that define responsibilities, monitoring the supplier’s performance through metrics (e.g., on-time delivery, deviation rates, quality of incoming material), and performing periodic raw material testing to verify the supplier’s Certificate of Analysis (CoA). You must demonstrate ongoing oversight, not just a one-time qualification.
With the revision of Annex 1, what is the single biggest cGMP shift in sterile manufacturing?
The biggest shift is the mandate for a formal, holistic Contamination Control Strategy (CCS). This is not just a collection of SOPs but a single, comprehensive document that justifies your facility design, processes, and monitoring programs based on risk management. It forces you to demonstrate how all your individual control measures (from gowning to HVAC to process design) work together to prevent contamination.
Why is Process Analytical Technology (PAT) considered a pillar of modern cGMP?
Because PAT embodies the core cGMP principle of building quality in, rather than testing it in. By providing real-time, in-process data, PAT allows for the active control of Critical Process Parameters (CPPs) to ensure Critical Quality Attributes (CQAs) are met. This shifts manufacturing from a rigid, recipe-based approach to a flexible, science-based model that can adjust to minor variabilities and guarantee a consistent outcome.
How should “human error” be treated as a root cause in a cGMP environment?
In a mature cGMP system, “human error” is rarely an acceptable root cause. It is usually a symptom of a flawed process or system. When an error occurs, the investigation must dig deeper: Was the procedure confusing? Was the training inadequate? Was the workspace poorly designed (human factors engineering)? Was the operator fatigued due to excessive overtime? A robust CAPA will address the underlying system failure, not just retrain the individual.
The Annual Product Review (PQR) is often seen as a chore. What is its intended cGMP purpose?
Its intended purpose is to be a proactive tool for continuous improvement. The PQR should not just be a retrospective data dump. It is a formal opportunity to analyze a year’s worth of data (trends, deviations, changes, stability results) to assess the health and consistency of a process. Its most important output should be a list of recommended CAPAs and process improvements for the upcoming year.
Related Readings
External Links on
국제 표준
- ISO 22716:2007 Good Manufacturing Practices (GMP) - Guidelines on Good Manufacturing Practices
- FDA 21 CFR Part 210 and 211: Current Good Manufacturing Practice in Manufacturing, Processing, Packing, or Holding of Drugs
- ISO 13485:2016 Medical devices - Quality management systems - Requirements for regulatory purposes
(링크 위에 마우스를 올려놓으면 콘텐츠에 대한 설명을 볼 수 있습니다.)
사용된 용어집
American Society for Testing and Materials (ASTM): an international standards organization that develops and publishes voluntary consensus technical standards for materials, products, systems, and services, aimed at improving quality and safety across various industries.
Application Programming Interface (API): 서로 다른 소프트웨어 애플리케이션이 통신하고 상호 작용할 수 있도록 하는 규칙 및 프로토콜 집합으로, 시스템 간 기능 통합 및 데이터 교환을 가능하게 합니다.
Attributable, Legible, Contemporaneous, Original, Accurate (ALCOA+): a set of principles ensuring data integrity in regulated environments, emphasizing that data must be traceable to its source, easily readable, recorded at the time of occurrence, in its original form, and precise in content.
Carbon Capture & Sequestration (CCS): a process that captures carbon dioxide emissions from sources like power plants and industrial processes, transporting it for storage underground or utilizing it in various applications, thereby reducing greenhouse gas concentrations in the atmosphere.
Certificate of Analysis (CoA): a document issued by a manufacturer or testing laboratory that confirms a product's specifications, quality, and compliance with regulatory standards, detailing test results and methods used for analysis.
Code of Federal Regulations (CFR): 미국 연방 기관에서 발행한 일반적이고 영구적인 규칙들을 주제별로 50개 편으로 정리한 것으로, 연방 규정에 대한 공식적인 법적 자료 역할을 합니다.
Contamination Control Strategy (CCS): a systematic approach to prevent, detect, and mitigate contamination in controlled environments, ensuring product quality and safety through defined procedures, monitoring, and risk management practices.
Corrective Action and Preventative Action (CAPA): a systematic approach to identifying, investigating, and addressing nonconformities and potential issues to prevent recurrence and ensure compliance with regulatory standards in quality management systems.
Corrective and Preventive Action (CAPA): 규제 기준 준수 및 프로세스와 제품의 지속적인 개선을 보장하기 위해 부적합 사항을 식별, 조사 및 해결하는 체계적인 접근 방식입니다.
Cost and Freight (CFR): 판매자가 지정된 항구까지 상품을 운송하는 데 드는 운송비와 운임을 부담하고, 구매자는 상품이 선박에 선적된 후 발생하는 보험 및 비용에 대한 위험과 책임을 부담하는 선적 계약.
Critical Control Points (CCP): specific stages in a process where control can be applied to prevent, eliminate, or reduce food safety hazards to acceptable levels. Identifying these points is essential for effective hazard analysis and critical control management in food production systems.
current Good Manufacturing Practice (cGMP): a system ensuring that products are consistently produced and controlled according to quality standards, encompassing regulations and guidelines for manufacturing processes, facilities, equipment, and personnel to ensure safety, quality, and efficacy in pharmaceuticals, food, and other regulated industries.
Design for Manufacturability (DFM): 제조 공정을 단순화하고 최적화하여 생산 비용을 절감하고 품질을 향상시키며 제품 출시 기간을 최소화하기 위해 설계 단계에서 제조 능력과 제약 조건을 고려하는 것을 목표로 하는 일련의 엔지니어링 관행입니다.
Device History Record (DHR): a compilation of records that documents the production history of a medical device, including manufacturing, quality control, and testing data, ensuring compliance with regulatory requirements and facilitating traceability throughout the device's lifecycle.
Device Master Record (DMR): a compilation of documents and specifications that provide the necessary information to produce a medical device, including design specifications, production processes, quality assurance measures, and labeling requirements, ensuring compliance with regulatory standards.
Failure Mode and Effects Analysis (FMEA): a systematic method for evaluating potential failure modes within a system, process, or product, assessing their effects on performance, and prioritizing risks to improve reliability and safety through corrective actions.
Food and Drug Administration (FDA): a federal agency of the United States Department of Health and Human Services responsible for regulating food safety, pharmaceuticals, medical devices, cosmetics, and tobacco products to ensure public health and safety through scientific evaluation and enforcement of compliance standards.
Good Manufacturing Practice (GMP): a system ensuring products are consistently produced and controlled according to quality standards, minimizing risks involved in pharmaceutical production and related industries. It encompasses guidelines for manufacturing processes, facility conditions, personnel qualifications, and documentation practices to ensure product safety and efficacy.
Hazard Analysis and Critical Control Points (HACCP): a systematic approach to food safety that identifies, evaluates, and controls hazards at critical points in the production process to prevent foodborne illnesses and ensure product safety.
Heating Ventilation and Air Conditioning (HVAC): a system designed to regulate indoor climate by controlling temperature, humidity, and air quality through heating, cooling, and ventilation processes. It includes components such as furnaces, air conditioners, ductwork, and thermostats for efficient environmental management.
Heating, Ventilation, and Air Conditioning (HVAC): 난방, 냉방 및 환기 과정을 통해 온도, 습도 및 공기 질을 제어하여 실내 환경을 조절하도록 설계된 시스템으로, 주거, 상업 및 산업 환경에서 쾌적함과 안전을 보장합니다.
High-Performance Liquid Chromatography (HPLC): 고압 하에서 고체 흡착 물질로 채워진 컬럼을 통해 액체 시료를 통과시켜 혼합물의 구성 성분을 분리, 식별 및 정량화하는 기술로, 시료와 고정상 사이의 상호 작용 차이를 이용합니다.
Installation Qualification (IQ): a documented process to verify that equipment or systems are installed according to specifications, including assessment of utilities, environmental conditions, and compliance with design requirements, ensuring readiness for operational qualification.
International Organization for Standardization (ISO): a non-governmental international body that develops and publishes standards to ensure quality, safety, efficiency, and interoperability across various industries and sectors, facilitating global trade and cooperation. Established in 1947, it comprises national standardization organizations from member countries.
Key Performance Indicator (KPI): a measurable value that demonstrates how effectively an organization is achieving key business objectives, often used to evaluate success at reaching targets.
Magnetic Resonance Imaging (MRI): a medical imaging technique that uses strong magnetic fields and radio waves to generate detailed images of internal body structures, particularly soft tissues, by detecting the signals emitted from hydrogen nuclei in the presence of a magnetic field.
Operational Qualification (OQ): a validation process that ensures equipment or systems operate according to specified requirements within defined limits, confirming that they perform as intended in their operational environment.
Partial Differential Equation (PDE): 미분방정식은 여러 개의 독립 변수, 미지의 함수, 그리고 그 편미분을 포함하는 수학 방정식입니다. 열전도, 유체 흐름, 파동 전파와 같은 현상을 설명하며, 해를 구하기 위해서는 특정한 경계 조건과 초기 조건이 필요한 경우가 많습니다.
parts per million (ppm): a unit of measurement representing the concentration of one substance in one million parts of another, often used to quantify pollutants or contaminants in air, water, or soil. It is equivalent to milligrams of substance per liter of solution or per kilogram of material.
Performance Qualification (PQ): a process that verifies a system or equipment operates according to specified requirements under real-world conditions, ensuring it consistently performs its intended function within predetermined limits.
Product Lifecycle Management (PLM): a systematic approach to managing a product's lifecycle from inception, through engineering design and manufacturing, to service and disposal, integrating people, processes, data, and technology to improve product quality, reduce time to market, and enhance collaboration across stakeholders.
Programmable Logic Controller (PLC): 산업 공정 자동화에 사용되는 디지털 컴퓨터로, 명령어를 저장하는 프로그래밍 가능한 메모리, 기계를 모니터링하고 제어하기 위한 입출력 인터페이스, 그리고 제어 알고리즘을 실행하는 프로세서로 구성됩니다.
Qualified Person (QP): an individual with the necessary education, experience, and authority to oversee and ensure compliance with regulatory requirements in the preparation and submission of technical documents, particularly in the mining and resource sectors, as defined by relevant industry standards.
Root Cause Analysis (RCA): 문제나 실패로 이어지는 근본적인 요인을 체계적으로 파악하는 과정으로, 단순히 증상만 치료하는 것이 아니라 근본적인 문제를 해결하고 제거하는 것을 목표로 합니다.
Standard Operating Procedure (SOP): a set of step-by-step instructions created to help workers carry out routine operations consistently and efficiently, ensuring compliance with regulations and quality standards.
Statistical Process Control (SPC): a method of quality control that employs statistical techniques to monitor and control a process, ensuring it operates at its full potential by identifying variations and maintaining consistent output within specified limits.