오염 제어 전략(CCS)은 모든 형태의 오염으로부터 제품 품질에 미치는 위험을 관리하기 위한 공식적인 접근 방식을 수립하는 포괄적이고 문서화된 프레임워크입니다. 이는 시설 설계 그 이상을 아우르는, 인력 교육, 물질 이동 프로토콜, 공정 흐름, 환경 모니터링, 설비 검증 등 다양한 요소를 하나의 일관된 계획으로 통합하는 살아있는 문서입니다. CCS는 기존 절차의 단순한 요약이 아니라, 위험 관리에 기반한 전략적 근거로서 특정 제어 조치가 시행되는 이유, 모니터링 방법, 그리고 의도된 제어 상태에서 벗어났을 때 취해야 할 조치를 명확히 정의합니다.
The cleanroom is one of the physical components, but it is fundamentally governed by the CCS. Its classification, operational parameters, and monitoring limits are direct outputs of the risk assessments detailed within the strategy. The CCS provides the rationale for the cleanroom’s existence and dictates its performance requirements to mitigate identified contamination risks.
핵심 요약
- CCS should be built upon the principles of Quality Risk Management (QRM)
- Holistic, facility-wide implementation strategy.
- Define and monitor critical control points.
- Personnel are a primary contamination source.
- Proactive and continuous environmental monitoring.
- Implement strict material and waste controls.
- Process design to minimize exposure
- 규정 준수를 보장합니다(예: 부록 1).
What is a Contamination Control Strategy (CCS)?
A Contamination Control Strategy (CCS) is a formal, comprehensive, and living document that defines a manufacturer’s entire approach to minimizing contamination in their products.
Mandated by the revised EU GMP Annex 1, its core purpose is to move away from a fragmented collection of separate procedures (for cleaning, gowning, monitoring, etc.) towards a single, holistic strategy that demonstrates a deep understanding of all potential contamination risks and the scientific rationale for the controls put in place to mitigate them.
The philosophy behind the CCS is that sterility and product quality are not achieved by any single action, but by the cumulative effect of a series of interconnected controls. The CCS is the master document that identifies every potential source of contamination—particulate, microbial, endotoxin/pyrogen, and chemical—and describes how the combination of facility design, equipment, procedures, and monitoring programs work together to protect the product throughout its lifecycle.
Key Elements of a Contamination Control Strategy
1. Facility and Equipment Design
This element is the physical foundation of the Contamination Control Strategy. It’s not enough to have a cleanroom; you ~ 해야 하다 justify why it was designed the way it was, based on risk.
Rationale for the design: the CCS must detail the logic behind the facility’s layout, including unidirectional flows for personnel, materials, equipment, and waste to prevent mix-ups and backtracking from “dirty” to “clean” areas. It must scientifically justify the 압력 cascades between rooms (e.g., the Grade B cleanroom is maintained at a significant positive pressure relative to the surrounding Grade C support area), providing data from the Building Management System (BMS) to prove these differentials are continuously maintained. The design of material and personnel airlocks, including their interlock mechanisms and purge times, must be rationalized as critical control points.
Equipment design: the Contamination Control Strategy justifies the selection of process equipment based on its ability to mitigate contamination. This includes specifying sanitary design features like crevice-free surfaces, 316L stainless steel, and Tri-Clamp fittings to prevent microbial colonization and facilitate effective cleaning. Crucially, it details the rationale for using advanced aseptic technologies. For example, it would justify the use of a Restricted Access Barrier System (RABS) or a fully contained isolator by referencing a risk assessment (FMEA) that shows these technologies significantly reduce the risk of operator-borne contamination compared to a conventional open-process cleanroom.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1: “Manufacture of Sterile Medicinal Products” (August 2022), specifically Section 4, “Premises.”
- FDA: Code of Federal Regulations, Title 21, Part 211.42, “Design and construction features” and 211.46, “Ventilation, air filtration, air heating and cooling.”
팁: use Computational Fluid Dynamics (CFD) modeling during the design phase of a new cleanroom or the installation of new equipment. Supplement this with physical smoke studies during qualification. The visual evidence from these studies provides irrefutable, science-based proof that your airflow patterns are unidirectional, effectively protecting the critical zone from contamination, which is far more powerful during an audit than simply presenting air velocity and pressure differential data.
2. Personnel
This element recognizes that humans are the primary source of microbial and particulate contamination in a cleanroom.
Gowning: 오염 제어 전략은 절차뿐 아니라 전체 가운 착용 시스템에 대한 세부 사항을 포함합니다. 여기에는 선택된 의복의 재질 과학(예: 보풀 방지, 액체 저항성), 멸균 주기 검증, 그리고 각 작업자에 대한 가운 착용 적합성 연구 결과가 포함됩니다. 이러한 연구는 개인이 가운의 멸균 상태를 손상시키지 않고 착용할 수 있음을 입증하는 객관적인 데이터(예: 장갑과 소매의 접촉면)를 제공해야 합니다.
Training and aseptic technique: 전략은 단순한 절차적 훈련 그 이상입니다.
Regulatory References:
- EU: 이는 운영자가 배지 채우기(무균 공정 시뮬레이션)에 성공적으로 참여함으로써 무균 조작에 대한 숙련도를 입증해야 하는 공식 자격 프로그램을 설명합니다. CCS는 규칙 이면의 "이유"를 강조하여 직원이 난기류 발생을 방지하기 위한 느리고 신중한 움직임의 중요성과 같은 오염 제어의 미생물학적 원리를 이해할 수 있도록 보장합니다.
- FDA: 연방 규정집, 제21권, 제211.28조, '인사 책임' 및 제211.113조, '미생물 오염 관리'
팁: implement a “human factors” study for your most critical aseptic interventions. Instead of just writing a procedure, observe operators performing the task and analyze the 인간 공학 and cognitive load. A procedure that is difficult, awkward, or confusing is a contamination risk. Simplifying the process based on human factors engineering is a proactive control that is far more effective than simply retraining personnel on a flawed procedure.
3. Utilities
Utilities that contact the product or product-contact surfaces are a direct potential pathway for contamination.
Control of critical utilities: the Contamination Control Strategy provides a comprehensive overview of the design, validation, and ongoing monitoring of critical utility systems. For a Water-for-Injection (WFI) system, it would detail the continuous hot-loop circulation (>80°C) to prevent biofilm formation, the validation of the multi-stage purification process, and the routine monitoring program for bioburden, endotoxin, Total Organic Carbon (TOC), and conductivity. Similarly, it would describe the filtration (e.g., sterile 0.22 µm filters) and qualification of process gases like compressed air or nitrogen that come into direct product contact.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 6, “Utilities.”
- FDA: Code of Federal Regulations, Title 21, Part 211.63, “Equipment design, size, and location,” supported by the principles in the “Guidance for Industry: Sterile Drug Products Produced by Aseptic Processing — Current Good Manufacturing Practice.”
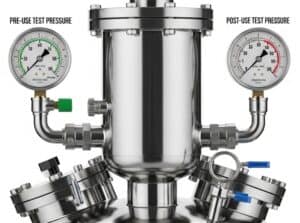
Application example: the CCS for a biologics facility details the risk assessment for the process air system. It identifies the point-of-use sterile filters on the bioreactor air inlet as Critical Control Points. The strategy can mandate an integrity test of these filters not only before the batch (pre-use) but also after the batch is harvested (post-use, pre-sterilization) to prove the filter remained integral throughout the entire cell culture process, thus ensuring the batch was protected.
팁: use online Total Organic Carbon (TOC) monitoring in your WFI loop not just as a quality attribute test, but as a leading indicator for system health. Establish a tight internal alert limit, well below the official action limit. A slow, upward trend in TOC levels is often the very first sign of developing biofilm in the system, allowing maintenance to intervene with sanitization cycles long before a significant microbial or endotoxin event occurs.
4. Raw Materials and Components
Contamination control begins with the materials that enter the facility.
Bioburden and endotoxin control: the CCS outlines a proactive strategy for managing the microbial quality of all incoming materials. This involves setting specifications for bioburden and endotoxin levels on raw materials, primary packaging (vials, stoppers), and product-contact components. This data is critical, as it is used to validate the robustness of downstream removal or sterilization steps (e.g., proving an autoclave cycle can achieve the required Sterility Assurance Level based on the maximum potential starting bioburden).
Supplier management: this section details the risk-based supplier qualification program. It describes how suppliers of critical components are audited and qualified, and the role of formal Quality Agreements in defining quality responsibilities. It also specifies the program for periodically verifying a supplier’s Certificate of Analysis (CoA) through in-house testing to ensure ongoing consistency and control.
Regulatory References:
- EU: EudraLex – Volume 4 – Part I, Chapter 5, “Production” and Chapter 7, “Outsourced Activities.”
- FDA: Code of Federal Regulations, Title 21, Part 211.84, “Testing and approval or rejection of components, drug product containers, and closures.”
팁: implement a risk-based program for your incoming component testing that goes beyond the standard pharmacopeia. For stoppers used in sensitive biologic drug products, your CCS should include a testing regime for extractables and leachables under worst-case conditions to ensure that no compounds that could degrade the drug molecule are introduced from the component itself.
Bioburden Details
The bioburden refers to the population of viable microorganisms present on a surface, in a liquid, or within a raw material before it undergoes a sterilization or disinfection process.
This microbial load, measured in Colony Forming Units (CFU), is a key indicator of the cleanliness of the manufacturing environment and a crucial factor in ensuring product safety and efficacy.
The primary goal in these industries is to maintain a low bioburden to minimize the challenge to the final sterilization process and to prevent product contamination. High bioburden levels can compromise the sterility of a product, potentially leading to infections in patients or defects in sensitive electronic components.
제약 산업의 바이오버든 The pharmaceutical industry operates under stringent regulatory requirements to control microbial contamination. The United States Pharmacopeia (USP) provides specific guidelines for bioburden limits in both sterile and non-sterile drug products. Non-sterile products: for non-sterile pharmaceuticals, the acceptable bioburden levels vary depending on the product’s intended use and route of administration. USP chapter <61> outlines the microbial enumeration tests, while chapter <1111> provides the acceptance criteria. These criteria are designed to ensure that the product is safe for its intended use and that the microorganisms present will not proliferate to harmful levels. Sterile products: for sterile drug products, the bioburden must be meticulously controlled before the final sterilization process. According to USP chapter <62>, sterile products should have a total aerobic microbial count (TAMC) and total yeast and mold count (TYMC) of no more than 10 CFU per 100 mL. The European Medicines Agency (EMA) also recommends a pre-sterilization bioburden limit of less than 100 CFU per 100 mL. | Bioburden in the medical device industry Similar to pharmaceuticals, the medical device industry places a strong emphasis on controlling bioburden to ensure the effectiveness of sterilization and the safety of the device. The international standard ISO 11737-1 provides guidance on the determination of the population of microorganisms on medical devices. The acceptable bioburden for a medical device can vary depending on its classification, intended use, and the materials it is made from. While there are no universal, mandated limits for all devices, a typical bioburden level for a medical device is between 0 and 150 CFUs. For some devices, a lower limit of less than 100 CFU may be targeted to ensure a higher sterility assurance level. The size and complexity of the device can also influence its bioburden, with larger and more intricate devices potentially having higher microbial loads. |
5. Cleaning and Disinfection
This element describes the chemical and physical removal of contaminants from surfaces (cleaning) and living organisms removal (disinfection).
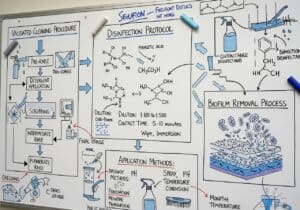
Validated procedures: the CCS provides the scientific justification for the entire disinfection program. This includes the rationale for selecting specific disinfectants (e.g., a broad-spectrum bactericide) and sporicides (e.g., a peracetic acid-based agent), supported by coupon studies that prove their efficacy against the facility’s common microbial isolates. It details the validated parameters for their use, including dilution, contact time, and application method.
Efficacy and rotation: the strategy must include a disinfectant rotation program to prevent the development of resistant microbial strains. The CCS explains the frequency and logic of this rotation. It also describes the validation of cleaning procedures for product-contact equipment, proving they can effectively remove both chemical residues of the previous product and any microbial contamination to pre-defined, health-based limits.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 5, “Cleaning and disinfection.”
- FDA: Code of Federal Regulations, Title 21, Part 211.67, “Equipment cleaning and maintenance.”
팁: your disinfectant efficacy validation should not be a one-time event. Your Contamination Control Strategy should mandate an ongoing “in-situ” efficacy program. Periodically, take environmental isolates (the actual bacteria and molds growing in your facility) and re-challenge your validated disinfectants in the lab. This verifies that your program remains effective against the current, relevant microflora, which can change over time.
6. Process Risk Management
This is the intellectual core of the CCS, where all potential hazards are formally identified and controlled.
Risk identification: the CCS must contain or reference a formal process risk assessment, typically an FMEA or HACCP. This assessment systematically breaks down the entire manufacturing process, step-by-step, to identify every potential contamination risk (e.g., aseptic connection, operator intervention, material transfer).
Critical Control Points: based on the risk assessment, the CCS identifies the Critical Control Points (CCPs)—the specific steps where control is essential to prevent or eliminate a risk. For each CCP, the CCS defines the specific control measure (e.g., use of a single-use sterile connector, continuous particle monitoring during filling) and the scientific rationale for why that control is effective.
Quality risk management always comes back to the risk benefit ratio of each control measure: a stricter cleanroom practice is justified only when its protection outweighs its cost. 팀 that document the risk to benefit ratio of every control can defend their choices during an audit.
Regulatory References:
- EU: EudraLex – 4권 – 부록 1(2022년 8월), 섹션 2, “의약품 품질 시스템(PQS)”, 품질 위험 관리(ICH Q9) 적용을 명시적으로 요구합니다.
- FDA: The principles are embedded throughout the Aseptic Processing Guidance, which is built on a foundation of risk management. The FDA has formally adopted the ICH Q9 guideline.
팁: make your process risk assessments “living documents.” The CCS should mandate a formal review of the FMEA for a given process after a set period (e.g., one year) or after a set number of batches. During this review, use actual deviation and batch failure data to re-score the “Probability” and “Detection” elements of the FMEA. This transforms the FMEA from a theoretical exercise into a dynamic, data-driven tool that accurately reflects the real-world performance of your process.
7. Environmental and Process Monitoring
This element describes the systems used to verify that the facility and process remain in a state of control.
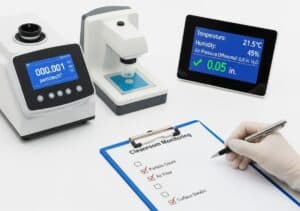
The EM program: the CCS provides a detailed rationale for the Environmental Monitoring (EM) program. It justifies every sampling location based on the process risk assessment (e.g., locations where sterile product is exposed, areas of frequent operator activity). It defines the methods (e.g., active air sampling, settle plates, contact plates), frequencies, and alert/action limits for both non-viable particulates and viable microbial counts.
Process monitoring: this section describes the use of modern monitoring technologies. This includes continuous, real-time particle monitoring in the Grade A critical zone, with alarms linked to the Batch Record. It also describes the trending of pressure differentials and other critical parameters. The Contamination Control Strategy must also define the program for identifying microbial isolates recovered from the EM program to the species level, which is crucial for tracking the facility’s microbial flora and investigating deviations.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 9, “Viable and non-viable environmental and process monitoring.”
- FDA: Code of Federal Regulations, Title 21, Part 211.113(b), which requires “appropriate written procedures… designed to prevent microbiological contamination.” The Aseptic Processing Guidance provides detailed expectations.
팁: use the microbial identification data from your EM program to create a “microbial map” of your facility. Track not just the counts, but the species and where they are found. This allows you to identify resident “house” organisms and track their movement. If a media fill or sterility test failure occurs, you can compare the contaminating organism to your facility map, which can dramatically accelerate the root cause investigation by pointing to the likely source area.
8. Continuous Improvement
The CCS is not a static document; it must evolve and is highly related to the “c” of the cGMP.
Data trending: the CCS mandates a formal system for the routine trending of all data related to contamination control, including EM results, personnel monitoring, utility monitoring, and batch-related deviations. Statistical Process Control (SPC) should be used to identify negative trends, even if the results are still within specification.
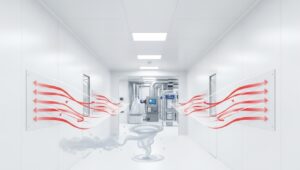
Feedback loop: this is the most critical part of the continuous improvement cycle. The Contamination Control Strategy must be a formal agenda item in the site’s Quality Management Review. The trend data must be reviewed by senior management, and the output must be a set of documented actions for improvement. If a new microbial strain is frequently recovered, for example, the CCS must be updated to reflect a potential change in the disinfection program or an investigation into its source.
Regulatory References:
- EU: EudraLex – 제4권 – 부록 1(2022년 8월), 섹션 2.5에는 CCS가 "적극적으로 업데이트되어야 하며 지속적인 개선을 추진해야 한다"고 명시되어 있습니다. 이는 ICH와 직접 연결됩니다. Q10, “Pharmaceutical Quality System.”
- FDA: The principles of continuous improvement are central to the PQS model described in ICH Q10, which the FDA has adopted.
팁: integrate the CCS review directly with the site’s Quality Management Review (QMR) and use a “Quality Risk Management” dashboard. Present KPIs derived from the CCS (e.g., EM excursion rates, media fill pass rates, contamination-related deviation trends) directly to senior leadership. This ensures that the health of your contamination control program has high-level visibility, which is critical for securing the resources and management commitment needed to drive meaningful continuous improvement.
클린룸
클린룸은 물류 절차 및 공급업체 관리와 함께 오염 제어 전략(CCS)의 중요한 부분입니다.
부인 성명: 다음 장들은 설계, 품질 및 공정 담당자들이 클린룸 원칙을 더 잘 적용할 수 있도록 구성되었습니다. 클린룸 관리는 매우 상세하지만, 다양한 제품과 요구 사항으로 인해 여러 요소와 특수성을 고려해야 하므로, 전문 인력과 공급업체의 도움이 필수적입니다.
클린룸 분류
이 표는 국제 표준에 따른 가장 일반적인 클린룸 등급을 비교합니다. ISO 14644-1 본 표준 및 유럽 연합의 EudraLex 4권 부록 1(GMP) 표준을 준수합니다.
ISO 클래스 5 EU GMP A등급 | ISO 클래스 6 | ISO 7등급 EU GMP B등급 | ISO 클래스 8 EU GMP 등급 C/D | |
| 공기 흐름 특성 | 단방향 흐름(UDF) 층류라고도 하는 이 흐름은 공기가 일정한 속도(일반적으로 0.36~0.54m/s)로 한 방향으로 이동하는 것을 말합니다. | 비단방향 흐름 난류 흐름이라고도 함: 오염은 희석을 통해 제어됩니다. | ||
| 주요 기능 | 중요 구역 보호 오염으로부터 노출된 멸균 제품 및 표면을 적극적으로 보호하도록 설계되었습니다. | 고품질 공정 제어 입자 수가 매우 적어야 하지만 완전한 무균 조건이 반드시 필요하지는 않은 공정에 사용됩니다. | 무균 공정 지원 A등급 구역의 즉각적인 배경 환경 역할을 합니다. | 일반 청소 지원 중요도가 낮은 활동의 배경으로 사용되거나, 상위 학년 학생들의 입장/가운 착용 장소로 사용됩니다. |
최대 입자 농도 (운영 중) | ISO 5:
A등급:
|
| ISO 7:
B등급:
| ISO 8 / C등급:
D등급: 작동상 정의되지 않음. |
최대 입자 농도 (휴식 중) | ISO 5 / A등급:
|
| ISO 7 / B등급:
| ISO 8 / C등급:
D등급:
|
미생물 한계 (운영 중) | Air Sample: <1 CFU/m³ Settle Plate (4 hrs): <1 CFU Contact Plate: <1 CFU | ISO에서 공식적으로 정의한 사항은 아니며, 적용 분야에 따라 다릅니다. | 공기 시료: 10 CFU/m³ 배양 후 침전 시간(4시간): 5 CFU 접촉판: 5 CFU | C등급: 공기: 100, 안정: 50, 접촉: 25 D등급: 공기: 200, 안정: 100, 접촉: 50 |
| 일반적인 공기 교환 횟수/속도 | 속도: 0.36 – 0.54 m/s (90 ft/min ± 20%) (ACH는 UDF의 주요 지표가 아닙니다.) | 하지만: 150~240 | 하지만: 60 90 | 하지만: 20 45 |
| 필수 복장 | 완전 무균복 착용: 무균 처리된 개별 포장 후드, 고글, 마스크, 작업복, 덧신. 무균 장갑 두 켤레 착용; 안쪽 장갑은 소매 아래로 넣어 착용하고, 바깥쪽 장갑은 위로 씌워 착용. 바깥쪽 장갑은 정기적으로 소독해야 합니다. | 완전 가운 착용: 청결 처리된 (반드시 멸균 처리될 필요는 없는) 작업복, 후드, 부츠, 장갑 및 마스크. | 완전 무균 가운 착용: A등급과 동일합니다. 모든 물품은 무균 상태여야 합니다. | C등급: 전용 수술복 또는 작업복, 전용 신발/덧신, 헤어넷, 수염 가리개, 마스크, 장갑. D등급: 깨끗한 작업복, 전용 신발 또는 신발 덮개, 헤어넷, 수염 덮개. |
| 필수 하드웨어 | • 필터링: 단방향 흐름을 구현하기 위해 터미널 전체에 HEPA/ULPA 필터가 장착되어 있습니다. • 표면: 제약 등급의, 비다공성, 비발포성, 매끄러운 표면(샌드위치 패널, 에폭시 타설/용접 비닐 바닥재). • 시공: 모든 이음매는 완전히 덮고 밀봉 처리했습니다. 돌출된 부분이 없습니다. | • 필터링: 터미널 간 천장에 필터가 있습니다. • 표면: 단단하고, 다공성이 없으며, 매끄러운 표면. • 시공 방식: 코브형 바닥. | • 필터링: 천장에 HEPA 필터가 설치되어 있습니다. • 표면: 단단하고, 다공성이 없으며, 털 빠짐이 없고, 매끄러운 표면. • 구조: 인원/자재용 연동식 에어록; 눈에 잘 보이는 차압계. | • Filtering: HEPA filters (often in AHU or ducted). • 표면: 매끄럽고 세척이 용이하며 다공성이 없는 바닥 및 벽. • 구조: 구분된 가운 착용 구역/에어록; 눈에 잘 띄는 차압계. |
| 있으면 좋은 하드웨어 | • 경보 기능이 내장된 실시간 입자 계수기. • 생물학적 오염 제거를 위한 자동 분무 시스템. • 절연 장치용 통합 장갑 무결성 테스트 장치. • 벽/바닥 몰딩이 완벽하게 통합되어 있습니다. | • 연속 입자 모니터링 시스템. • HEPA 필터가 장착된 연동식 통과 챔버. • 내화학성 바닥재. | • 중요 구역 주변의 입자 실시간 모니터링. • 자동 고속 롤업 도어. • 벽면에 각종 설비가 내장된 패널로 깔끔하게 정리할 수 있습니다. | • 네트워크 연결형 입자 계수기. • 자동 신발 덮개 부착기. • 벽걸이형 스테인리스 스틸 가구. |
| 일반적인 적용 사례 | • 바이알의 무균 충전 • 무균 연결 및 이송 • 멸균 재료의 개방형 처리 • 세포 치료 처리 과정 | • 고급 의료기기 조립 • 비멸균 상태이지만 미생물 오염도가 낮은 의약품 충전 • 고품질 광학 부품 조립 | • A등급 구역을 위한 배경 공간 • 완충액 및 배지 준비 • 무균 부품 조립 | • C등급: 구성 요소 세척, 중요도가 낮은 무균 공정, B등급을 위한 기초 작업. • D등급: 가운 착용실, 중요도가 낮은 제형, 초기 준비 단계. |
| 준거기준 | 미국/국제: ISO 14644-1:2015 EU: EudraLex 제4권, 부록 1(2022) | 미국/국제: ISO 14644-1:2015 | 미국/국제: ISO 14644-1:2015 EU: EudraLex 제4권, 부록 1(2022) | 미국/국제: ISO 14644-1:2015 EU: EudraLex 제4권, 부록 1(2022) |
| 일반적인 오염물질 및 크기 | 이 기준은 특정 유형이 아닌 총 입자 수를 측정합니다. 그러나 주요 오염원과 그 일반적인 크기는 다음과 같습니다.
| |||
오염 비교
클린룸 기준의 엄격함을 이해하려면 일상 환경의 입자 수 제한과 비교해 보는 것이 도움이 됩니다. (이 값들은 일반적인 값이며, 물론 실제 값은 매우 다양할 수 있습니다.)
| 사무실 | 집의 방 | 주방 (요리 중) | 일반 공장 | |
| 일반적인 입자 수(m³당) | 약 1,900,000개 (≥0.5 µm) | >100,000,000 (≥0.3 µm) | 극도로 높은 급증세, 잠재적으로 850억 이상 | 변동성이 매우 크며, 종종 3,500만 이상입니다. |
| 일반적인 입자 크기 | 미크론 크기의 먼지와 섬유 | 알레르기 유발 물질 및 초미세 입자를 포함한 광범위한 범위 | Primarily ultrafine particles (<0.1 µm) and fine particles (PM2.5) | 공정에 따라 크기가 다른 먼지부터 미세한 서브마이크론 입자까지 다양합니다. |
| 1차 자료 | 먼지, 의류/카펫 섬유, 피부 각질, 냉난방 공조 장치 흡입구 | 먼지, 애완동물 비듬, 꽃가루, 곰팡이 포자, 인간 활동 | 연소, 조리(튀김, 구이) 시 발생하는 에어로졸 형태의 기름 및 지방 | 산업 공정(분쇄, 절단), 원자재, 연소 부산물 |
결론: 알레르기가 있다면 집에서 일하는 것보다 사무실에서 일하는 게 훨씬 낫죠 😉
최고의 클린룸: 그 “메모리 시설위대한 것의 블레이드 러너 2049 영화드니 빌뇌브 감독이 필립 K. 딕의 소설을 원작으로 제작했습니다.
클린룸 최적 설계 사례
제약에서부터 반도체 제조 분야에서 클린룸 설계 원칙은 공기 흐름 관리부터 건축 자재 선택에 이르기까지 모든 것을 포괄하는 다각적인 접근 방식에 기반을 두고 있습니다.
전략적 레이아웃 및 워크플로우 최적화오염 위험을 최소화하고 운영 효율성을 향상시키려면 잘 설계된 레이아웃이 매우 중요합니다. 설계는 적절한 절차를 시행하고 사람과 자재의 이동을 유도하여 교차 오염을 방지해야 합니다. 용도지역 및 인접성: 청결도가 점차 높아지는 구역을 명확하게 구분하여 레이아웃을 설계하십시오. 외부 구역에서 가운 착용실을 거쳐 최종적으로 메인 클린룸으로 이동하도록 하면 완충 지대가 형성되어 가장 중요한 구역을 보호하는 데 도움이 됩니다. 단방향 흐름: 교차 오염을 방지하기 위해 인력, 자재 및 폐기물의 일방향 흐름을 계획해야 합니다. 이는 가능한 경우 별도의 출입구를 설정하고 오염된 물질이 깨끗한 물질과 섞이지 않도록 하는 것을 의미합니다. | 세심한 HVAC 시스템 설계냉난방 및 환기(HVAC) 시스템은 클린룸의 핵심으로, 공기 청정도, 온도, 습도 및 압력을 유지하는 역할을 합니다. 이 시스템은 시설 전체 에너지 사용량의 50~80%를 차지할 수 있습니다. 에프여과 수준: 0.3미크론 크기의 입자를 99.97%까지 포집하는 고효율 미립자 공기(HEPA) 필터가 기본 사양입니다. 가장 엄격한 등급(ISO 5 이하)의 경우, 초저침투 공기(울파더 작은 입자까지 포착하려면 필터가 필요할 수 있습니다. 공기 흐름 패턴: 단방향(층류) 기류와 비단방향(난류) 기류 중 어떤 것을 선택할지는 요구되는 ISO 등급에 따라 달라집니다. 공기가 한 번에 한 방향(일반적으로 아래쪽)으로 흐르는 단방향 기류는 ISO 5와 같은 엄격한 등급에서 중요 공정으로부터 입자를 제거하는 데 사용되는 반면, 난류는 덜 중요한 환경(ISO 6 이상)에 적합합니다. |
오염 제어를 우선시하십시오효과적인 오염 제어 전략은 최우선 과제이며 설계 초기 단계부터 통합되어야 합니다. 이는 인력, 자재, 장비를 포함한 모든 잠재적 오염원을 고려하는 총체적인 접근 방식을 의미합니다. 에어 샤워: 매우 민감한 클린룸의 경우, 가운 착용 구역 입구에 에어 샤워를 설치할 수 있습니다. 고속 HEPA 필터로 여과된 공기 분사기가 작업자가 청정 공간으로 들어가기 전에 몸에 묻은 미세 입자 오염 물질을 제거합니다. 엄격한 가운 착용 규정: 설계는 엄격한 가운 착용 절차를 지원해야 합니다. 여기에는 작업자가 오염물질이 떨어지지 않는 클린룸 의복을 올바른 순서로 착용할 수 있도록 충분한 공간과 시설을 제공하는 것이 포함됩니다. |  |
재료의 신중한 선택건축에 사용되는 자재는 털이 빠지지 않고, 다공성이 없으며, 청소가 용이해야 할 뿐만 아니라 세척제와 소독제에 대한 저항성도 있어야 합니다. 일반적인 재료 선택: 스테인리스강은 내구성, 매끄러운 표면, 부식 및 화학 물질에 대한 저항성 때문에 선호되는 경우가 많습니다. 그 외 벽 패널에는 에폭시 코팅 표면, 유리섬유 강화 플라스틱(FRP), 코팅 알루미늄 등이 일반적으로 사용됩니다. 바닥재 선택 시 고려 사항: 바닥재는 이음매 없이 시공 가능하고 내구성이 뛰어나야 합니다. 비닐 및 에폭시 바닥재는 이음매 없이 시공할 수 있어 먼지 축적 위험을 줄이고, 전자 제품 제조에 필수적인 정전기 방지 기능을 제공하기 때문에 널리 사용됩니다. | 이음매 없고 밀봉된 표면입자 축적을 방지하고 효과적인 청소를 용이하게 하기 위해 벽, 바닥, 천장의 모든 이음새, 틈새 및 관통부는 적절하게 밀봉되어야 합니다. 곡선형 전환: 바닥과 벽이 만나는 부분, 그리고 벽과 천장이 만나는 부분에는 몰딩을 사용하세요. 이렇게 하면 곡선으로 매끄럽게 연결되어 청소가 쉬워지고 날카로운 모서리에 오염 물질이 쌓이는 것을 방지할 수 있습니다. 플러시 통합: 창문, 조명기구, 콘센트는 벽이나 천장 표면과 평평하게 설치되도록 설계해야 합니다. 이렇게 하면 먼지나 이물질이 쌓일 수 있는 틈새가 없어 청소가 더욱 효과적입니다. |
 | 제어 환경 매개변수온도와 습도를 정밀하게 제어하는 것은 여러 민감한 공정뿐만 아니라 작업자의 쾌적함과 오염 방지에도 매우 중요합니다. 습도의 이중적 위협: 높은 습도(일반적으로 60% 이상)는 미생물 번식을 촉진하고 민감한 장비의 부식을 유발할 수 있습니다. 반대로 낮은 습도(30% 미만)는 정전기 발생을 초래하여 전자 부품을 손상시키고 표면에 이물질이 달라붙게 할 수 있습니다. 온도 및 인력: 불편한 온도는 작업자가 땀을 흘리거나 떨게 하여 입자 방출량을 증가시키고 클린룸 의복의 효과를 저하시킬 수 있습니다. 일반적인 목표 온도는 21°C(70°F)이며 약간의 편차가 허용됩니다. (저희의 £8220도 참조하세요.)업계 비밀이지만, 누구도 자랑스러워하지 않는 비밀” 인 11. 장갑을 낀 손 루틴 아래는 실내 온도 조절과 연관되어 땀 발생을 최소화하며, 날카로운 모서리나 버(burr)가 없어야 합니다. |
양압 및 음압인접 지역 간의 적절한 기압 차이를 유지하는 것은 오염 물질의 유입을 방지하는 데 매우 중요합니다. 공기는 자연적으로 압력이 높은 곳에서 낮은 곳으로 흐릅니다. 연쇄적인 압력: 일반적인 전략은 가장 깨끗한 방의 압력을 높게 유지하는 계단식 압력 차이 방식입니다. 각 바깥쪽 방이나 에어록은 압력이 조금씩 낮아지도록 설계되어 공기가 항상 가장 깨끗한 공간에서 빠져나가도록 하고 오염 물질이 유입되는 것을 방지합니다. 기포형, 수축형, 계단식 에어록: 목적에 따라 다양한 압력 방식을 사용할 수 있습니다. '버블' 에어록은 주변 공간보다 높은 압력을 유지하여 에어록 내부를 보호합니다. '싱크' 에어록은 낮은 압력을 유지하여 오염 물질을 내부에 가둡니다. '캐스케이드' 에어록은 고압 클린룸과 저압 통로 사이의 중간 압력을 유지합니다. | 모듈식 및 유연한 디자인모듈형 요소를 설계에 통합하면 향후 확장, 이전 및 변화하는 프로세스 요구 사항에 대한 적응이 최소한의 중단으로 가능하지만, 신중하게 평가해야 합니다(아래 팁 참조). 공사 기간 단축: 구성 요소가 현장 외부에서 사전 제작되기 때문에 현장 조립 속도가 기존 건설 방식보다 훨씬 빠르며, 프로젝트 기간과 가동 중지 시간을 줄여줍니다. 확장성 및 재구성: 모듈형 클린룸은 공정 요구 사항이 변경될 경우 더 쉽게 확장하거나 재구성할 수 있습니다. 이러한 적응성 덕분에 모듈형 클린룸은 장기적으로 비용 효율적인 투자가 됩니다. 팁: 일반적으로 장비와 벤치에는 바퀴를 추천하지만 린 제조 생산의 모듈성을 개선함으로써 이를 최소화할 수 있습니다. 여기서는 바퀴나 브레이크에서 발생할 수 있는 입자 때문에 이를 최소화해야 합니다. |
통합형 및 클린룸 호환 조명조명은 클린룸의 무결성을 손상시키지 않으면서 세밀한 작업에 필요한 충분한 조도를 제공해야 합니다. 밀폐형 및 매립형 설비: 조명기구는 먼지나 이물질이 쌓이거나 떨어지는 것을 방지하기 위해 완전히 밀봉되어야 합니다. 또한 천장에 밀착되도록 설치해야 하며, 필요한 경우 덮개를 씌워야 합니다. '물방울' 모양공기 흐름 패턴을 방해하지 않도록 하기 위함입니다. |  |
세심하게 디자인된 드레스룸가운 착용실 또는 전실은 외부 환경과 클린룸 사이의 중요한 완충 지대 역할을 합니다. 효과적인 오염 제어를 위해서는 전실의 설계가 매우 중요합니다. 경계 및 흐름: 잘 설계된 가운 착용실은 '더러운' 구역과 '깨끗한' 구역을 명확하게 구분하는 경계선이 있으며, 종종 '앉아서 쉴 수 있는 벤치'가 포함되어 있습니다. 레이아웃은 직원이 논리적이고 순차적인 가운 착용 순서를 따르도록 안내해야 합니다. 적절한 편의 시설: 해당 공간에는 의복 보관 및 착용 과정을 용이하게 하기 위한 사물함, 장갑 및 마스크 디스펜서, 자가 검사용 거울과 같은 클린룸 전용 가구가 갖춰져 있어야 합니다. | 최소화된 가구 및 장비이것들은 린 원칙 그리고 5S하지만 이러한 원칙들을 최대한 적용해야 합니다! 오염 가능성을 줄이고 청소가 필요한 표면을 최소화하기 위해 클린룸 내부에는 필수적인 가구와 장비만 배치해야 합니다. 클린룸 등급 재료: 모든 품목은 스테인리스강과 같이 털이 빠지지 않고 다공성이 없는 재질로 만들어져야 합니다. 폴리프로필렌 쉽게 소독할 수 있는 것들. 인체공학적 그리고 미니멀리즘 디자인: 가구는 입자가 쌓일 수 있는 수평면을 최소화하도록 설계되어야 합니다. 또한 효율적이고 안전한 작업 방식을 지원할 수 있도록 인체공학적으로 설계되어야 합니다. 재고도 없고, 예비 부품도 없고, 설명서도 없다. |
 | 모니터링 시스템강력한 모니터링 시스템을 설치하여 주요 환경 매개변수를 지속적으로 추적 및 기록함으로써 지속적인 규정 준수를 보장하고 즉각적인 시정 조치를 취할 수 있도록 하십시오. 모니터링해야 할 주요 매개변수: 이 시스템들은 공기 중 입자 수, 기압 차이, 온도 및 습도를 실시간으로 추적합니다. 통제 vs. 모니터링: 설정값을 유지하기 위해 냉난방 시스템을 능동적으로 제어하는 건물 관리 시스템(BMS)과 품질 보증 및 규정 준수를 위해 데이터를 독립적으로 기록하고 매개변수가 사양을 벗어날 경우 경보를 울리는 환경 모니터링 시스템(EMS)을 구분하는 것이 중요합니다. |
전용 인원 출입용 에어록(PAL)인원 에어록(PAL)은 통제되지 않은 구역이나 저등급 구역과 클린룸 사이를 이동하는 직원을 위한 주요 차단막 역할을 하는 특수실입니다. 그 목적은 사람에 의해 유입되는 오염을 제어하는 것입니다. 서로 맞물린 문: PAL의 핵심적인 특징 중 하나는 출입문이 동시에 열리는 것을 방지하는 연동 도어 시스템입니다. 이는 두 공간 사이의 압력 차이를 유지하고 공기 중 오염 물질이 청정 공간으로 직접 유입되는 것을 막습니다. 가운 착용 및 가운 벗기 절차: PAL은 종종 가운 착용실과 동의어로 사용되며, 클린룸 의복을 올바른 순서로 착용하고 벗을 수 있도록 설계되었습니다. 설계는 사용자가 과정을 따라갈 수 있도록 물리적으로 안내하여 오류를 최소화해야 합니다. | 상품 및 자재의 입출고(MAL)자재 에어록(MAL)은 클린룸의 무결성을 손상시키지 않고 자재, 장비 및 완제품을 클린룸 안팎으로 이동시키기 위해 설계된 전용 공간입니다. 통과구 및 대형 에어록: 작은 물품의 경우, 잠금 장치가 있는 벽걸이형 통과형 보관함이 효율적인 해결책입니다. 더 큰 장비나 카트의 경우, 자재 보관을 위해 설계된 워크인형 자재 보관실(MAL)이 필요하며, 이는 일반 보관실(PAL)과 유사한 기능을 합니다. 오염 제거 및 닦아내기: 설계 시 클린룸에 반입하기 전에 자재를 소독할 수 있도록 해야 합니다. 일반적으로 이는 작업자가 적절한 멸균제를 사용하여 모든 품목을 닦을 수 있는 공간과 표면을 제공하는 것을 의미합니다. 일부 고급 MAL(다기능 의료기기)에는 기화 과산화수소(VHP) 분무기와 같은 자동 소독 시스템이 통합될 수 있습니다. |
에너지 효율성 및 지속가능성에 중점을 둡니다.
클린룸은 에너지 소비가 매우 높기 때문에, 에너지 효율을 높이기 위해 다음과 같은 요소들을 통합하는 것이 중요합니다. 지속 가능한 디자인 이러한 원칙들은 장기적인 운영 비용과 환경 영향을 크게 줄일 수 있습니다. 주요 요인은 단열과 가변 풍량(VAV) 시스템: VAV 시스템은 HVAC 시스템을 최대 출력으로 지속적으로 가동하는 대신, 입자 수나 재실률과 같은 실시간 조건에 따라 공기 흐름을 조절하여 활동이 적은 시간대의 에너지 소비를 줄일 수 있습니다.
팁: 업종상 방문객이 잦다면, 방문객을 직접 들여보내는 대신 큰 창문이나 유리 터널을 설치하는 것을 고려해 보세요. (이는 또한 여러분이 고객을 진심으로 배려하고 절차를 엄격하게 준수한다는 것을 보여줄 것입니다.)
추가 팁: 모든 자재가 매우 특수하기 때문에 기존 시설을 필사적으로 업그레이드하려고 애쓰는 것보다 벽과 천장을 포함하여 클린룸을 처음부터 새로 짓는 것이 비용이 덜 들 수 있습니다.
클린룸 인체공학적 모범 사례
위에서 설명한 바와 같이 뛰어난 클린룸 설계와 최첨단 장비는 무균 환경을 유지하기 위한 요소 중 일부일 뿐입니다.
클린룸에서 오염을 일으키는 가장 큰 원인은 바로 그 안에서 일하는 사람들입니다.
그러므로 특정한 인간의 관행과 행동에 대한 엄격한 준수는 권장 사항에 그치는 것이 아니라 절대적으로 필수적입니다.
1. 가운 착용 및 탈의 순서이것이 첫 번째 방어선입니다. 담당자는 클린룸 의복을 착용하고 벗는 정확하고 단계적인 절차를 숙지해야 합니다. '가장 깨끗한 것부터 가장 더러운 것까지' 원칙: 멸균복 착용은 멸균 의복의 오염을 최소화하는 순서를 따라야 하며, 일반적으로 머리부터 아래쪽으로 (후드, 마스크, 작업복) 착용하고 마지막에 장갑을 착용하는 것이 일반적입니다. 비접촉 기술: 멸균 장갑을 착용할 때는 장갑의 멸균된 바깥쪽 면이 맨손이나 멸균되지 않은 표면에 닿지 않도록 하는 방법을 반드시 사용해야 합니다. | 2. 의도적이고 통제된 움직임클린룸 내부의 모든 움직임은 입자의 생성 및 확산을 줄이기 위해 느리고 신중하며 최소화되어야 합니다. '클린룸 투어': 직원들은 팔을 흔들지 않고 천천히 일정한 속도로 걷도록 훈련받아야 합니다. 빠르고 갑작스러운 움직임은 표면에 가라앉은 입자를 휘저어 올릴 수 있는 난류를 발생시킵니다. 신체 움직임을 최소화하세요: 스트레칭, 긁기, 급격한 고개 돌리기와 같은 불필요한 동작은 피하십시오. 필요한 모든 동작은 효율적이고 부드럽게 수행해야 합니다. |
3. 철저한 개인위생오염 관리는 작업자가 탈의실에 들어가기 전부터 시작됩니다. 화장품 및 향수 사용 금지: 화장품, 헤어스프레이, 향수, 로션은 엄청난 양의 미세 입자와 휘발성 유기 화합물(VOC)을 방출할 수 있습니다. 이러한 제품들은 엄격히 금지되어야 합니다. 입장 전 손 씻기: 가운 착용 구역에 들어가기 직전에 손과 팔뚝은 지정된 항균 비누로 깨끗이 씻고 보풀이 없는 수건으로 즉시 말려야 합니다. |  |
4. 올바른 '클린룸 자세'개인이 서 있는 자세와 작업 공간과의 상호 작용 방식은 오염에 직접적인 영향을 미칩니다. 몸을 앞으로 기울이세요, 옆으로 넘기지 마세요: 작업대나 공정 스테이션에서 작업할 때는 허리를 숙여 상체를 기울이고 머리와 몸통을 작업 영역에서 최대한 멀리 떨어뜨려야 합니다. 제품이나 공정 위로 몸을 직접 숙이는 것은 절대 금지입니다. 공기 흐름을 존중하세요: 단방향(층류) 흐름 환경에서 작업자는 자신의 신체가 중요한 오염원이라는 점을 인지해야 합니다. 깨끗하고 여과된 공기가 제품에 먼저 도달하도록 작업자는 항상 중요 제품이나 공정의 하류에 위치해야 합니다. | 5. '퍼스트 에어' 마인드이 원칙은 단방향 공기 흐름 클린룸에서 가장 중요합니다. '최초 공기'는 어떤 물체를 통과하기 전에 HEPA/ULPA 필터를 직접 통과한 깨끗하고 여과된 공기입니다. N공기 흐름을 차단한 적이 있습니까? 작업자는 절대로 필터 표면과 중요 제품 사이에 물건, 손 또는 도구를 넣어서는 안 됩니다. 이렇게 하면 난류가 발생하여 오염될 가능성이 있는 '그림자' 형태의 공기 흐름이 생깁니다. 가장 깨끗한 구역에서 작업하세요: 모든 중요한 조작은 '첫 번째 공기'의 직접 경로, 즉 가능한 한 상류에서 수행되어야 합니다. |
 | 6. 노출된 피부에 대한 '접촉 금지' 정책적절한 보호복을 착용하더라도 피부가 우발적으로 노출될 수 있습니다. 얼굴과 복장 규정: 작업자는 클린룸 내부에 있는 동안 절대로 얼굴을 만지거나 마스크 또는 후드를 조정하거나 노출된 피부 부위를 만지지 않도록 교육받아야 합니다. 조정이 필요한 경우, 클린룸 밖으로 나가 다시 가운을 착용해야 합니다. 장갑의 내구성: 장갑은 클린룸용 소독제(예: 70% 이소프로필 알코올)로 자주 소독하고 찢어짐이나 구멍이 있는지 정기적으로 검사해야 합니다. |
7. 클린룸 의사소통 방법말하는 행위는 상당량의 입자와 물방울 오염을 발생시킵니다. 대화를 최소화하세요: verbal 의사소통 should be kept to an absolute minimum. 비언어적 신호와 시스템을 활용하세요: 가능한 한 손짓, 사내 전화 시스템 또는 화이트보드를 활용하여 의사소통하십시오. 말로 소통해야 할 경우에는 중요한 작업 과정에서 등을 돌린 자세로 이야기해야 합니다. | 8. 자재 및 폐기물 흐름 프로토콜자재를 다루는 방식은 사람들이 이동하는 방식만큼이나 중요합니다. '트리플 랩' 항목: 클린룸으로 자재를 반입할 때는 보통 3중 포장 시스템을 사용합니다. 가장 바깥쪽 포장은 자재 에어록에서 제거하고, 두 번째 포장은 가운 착용실에서 제거하며, 마지막 포장은 사용 시점에만 제거합니다(구성 요소 및 공급업체에 대한 자세한 내용은 아래 장을 참조하십시오). 전용 폐기물 수거: 폐기물은 지정된 클린룸 호환 용기에 담아 수거하고, 반출 과정에서 청정 구역을 오염시키지 않도록 특정 절차에 따라 처리해야 합니다. |
9. 개인 소지품공정에 필수적이지 않은 모든 것은 오염의 원인이 됩니다. 개인 소지품에 대한 무관용 원칙: 여기에는 장신구(장갑 착용 시에도 포함), 시계, 휴대폰, 열쇠, 그리고 클린룸 사용을 위해 특별히 설계되지 않은 모든 종이나 필기구가 포함됩니다. 승인된 클린룸 용품: 먼지가 날리지 않도록 설계된 클린룸 인증 용지, 펜 및 도구만 사용하십시오. |  |
10. 건강 및 복지 정책작업자의 건강은 클린룸 환경에 직접적인 영향을 미칩니다. 아플 땐 집에 머무르세요: 감기, 기침, 피부 질환과 같은 질병이 있을 경우 직원들이 이를 보고하고 집에 머물도록 장려하는 정책이 마련되어야 합니다. 아픈 몸으로 출근하는 것은 심각한 오염 위험을 초래합니다.  모든 피부 병변을 덮으십시오: 베인 상처, 찰과상 또는 일광 화상은 가운을 입기 전에 털이 빠지지 않는 승인된 붕대로 완전히 덮어야 합니다. | 11. 장갑을 낀 손 루틴장갑은 장비 및 재료와의 주요 접촉점이며 쉽게 교차 오염될 수 있습니다. 주요 작업 전후에 소독하십시오: 중요 제품을 취급하기 전, 중요하지 않은 표면(예: 장비 패널)을 만진 후, 그리고 정기적으로 장갑을 착용한 손에 70% IPA 멸균 용액을 뿌리고 자연 건조시켜야 합니다. 올바른 소독 방법: 두 장갑의 전체 표면이 소독제로 완전히 젖도록 하십시오. 업계의 비밀 (아무도 자랑스러워하지 않는): 장갑은 한 겹 또는 두 겹으로 착용하면 밀폐성이 높아 작업자가 땀을 흘리게 됩니다. 또한 일회용이고 얇기 때문에 쉽게 찢어집니다. 그렇다면 장갑이 찢어지면 땀은 어디로 갈까요? 장갑 교체 주기를 엄격하게 정하고, 땀 발생을 최소화하기 위해 작업장 온도도 조절해야 합니다. |
 | 12. 가운 착용실을 전환 구역으로 활용하기탈의실은 휴게실이 아닙니다. 중요 관리점. 사교 활동 금지: 가운 착용실은 가운을 입고 벗는 용도로만 사용해야 합니다. 그곳에서 사적인 대화를 나누어서는 안 됩니다. 여기엔 커피 머신이 없어요! 경계선을 따라가세요: '앉아서 쉴 수 있는 벤치' 또는 이와 유사한 구분대가 '오염된' 구역과 '깨끗한' 구역을 분리합니다. 직원은 '오염된' 쪽의 신발이나 의복이 '깨끗한' 쪽 바닥에 닿지 않도록 해야 합니다. |
팁: 제대로 복장을 갖춘 요원의 경우 눈만 보여야 하므로, 옷에 각자의 이름을 표시하는 것이 필요할 수 있습니다. 단, 벨크로는 사용하지 않아야 합니다. (이는 요원들이 가능한 한 눈에 띄지 않아야 하고, 서로를 잘 알아야 한다는 점과 균형을 이루어야 합니다.)
인사 행동, 자격 요건, 개인적 책임과 절차 준수
일반적으로 규제 산업의 모든 공정에서와 마찬가지로, 모든 직원은 기밀 구역에 출입하기 전에 정식 자격을 갖춰야 합니다. 자격은 해당 구역별 표준 작업 절차(SOP), 무균 행동 및 오염 제어 원칙을 다루는 문서화된 교육 프로그램을 통해 취득됩니다. 이 교육은 직원들이 항상 상황을 인지하고 자신의 작업 구역에 대한 특정 요구 사항을 이해하도록 하는 것을 목표로 합니다.
공급업체 사양
여기서는 클린룸용 제품을 공급하는 공급업체에게 제공하는 구체적인 지침의 몇 가지 전형적인 예만 자세히 설명합니다.
팁: 클린룸용 제품 공급 경험이 없는 공급업체라면 상당한 학습 곡선을 거쳐야 할 수도 있다는 점을 미리 알려주세요 (제 경험상, 라벨에 특수 재질이 필요하며 단순히 '붙이는 것'만으로는 충분하지 않다는 점을 설명하는 것부터 시작하면 됩니다).
참고: 일반적인 규제 산업 사양은 여기에 포함되지 않습니다(공급되는 자재에는 재료 구성, 로트 번호 및 관련 표준 준수 여부를 명시하는 적합성 인증서(CoC)와 분석 인증서(CoA)가 함께 제공되어야 합니다. 원자재에서 완제품까지 완전한 추적성이 요구되며, 관련 기록은 유지 관리되고 감사를 위해 제공되어야 합니다).
1. 필수 재료 구성 및 표면 마감
모든 구성 요소는 본질적으로 입자 발생량이 적고 가스 방출량이 적은 재료로 제작되어야 합니다.
- 금지 품목: 최종 부품에는 가공되지 않은 목재, 종이 기반 제품, 코팅되지 않은 탄소강, 천연 섬유 직물(예: 면, 양모) 또는 분말이나 실리콘을 함유한 재료가 절대 사용해서는 안 됩니다.
- 표면 요구 사항: 표면은 입자 탈락을 방지하고 청소를 용이하게 하기 위해 비다공성이고 매끄러워야 합니다. 금속의 경우 전해연마와 같은 마감 처리가 필요합니다. 플라스틱성형 과정에서 플래시와 버가 없는 매끄러운 표면 마감이 필수적입니다. 해당되는 경우 최대 표면 조도(예: Ra < 0.8 µm)를 지정하십시오.
2. 검증된 정밀 세척 공정
모든 제품은 포장 직전에 최종 검증된 세척 과정을 거쳐야 합니다.
- 방법론: 이는 일반적으로 여러 단계를 거칩니다. 초음파 클린룸 사용에 승인된 세제를 사용한 세척 공정 후, 가열된 고순도 탈이온수(DI)로 여러 차례 연속 헹굼을 진행합니다.
- 최종 헹굼 확인: 최종 탈이온수 헹굼액의 수질을 검사해야 합니다. 모든 잔류물과 이물질이 제거되었음을 입증하기 위해 최종 헹굼액이 저항률(예: >18 MΩ·cm), 총 유기 탄소(TOC) 및 액체 입자 수(LPC)에 대한 규격을 충족함을 보여주는 데이터를 제공해야 합니다.
- 건조 프로토콜: 부품은 가열된 HEPA 또는 ULPA 필터가 장착된 질소 또는 깨끗하고 건조한 공기를 불어넣는 것과 같은 오염 방지 방법을 사용하여 건조해야 합니다. 이 최종 단계에서는 '보풀이 적은' 물티슈를 포함하여 어떤 종류의 천으로도 닦는 것이 금지되어 있습니다.
3. 이중 포장 의무화
모든 품목은 인증된 클린룸 환경 내에서 이중 포장해야 합니다. 이 절차는 절대 타협할 수 없으며 정확하게 준수해야 합니다.

- 포장 환경: 내부 백은 최종 사용자의 클린룸과 동일하거나 더 깨끗한 환경(예: ISO 5 클린룸으로 들어가는 제품의 경우 ISO 5 환경)에서 밀봉되어야 합니다.
- 가방 소재: 새롭고 인증된 클린룸 등급의 제품만 사용하십시오. 폴리에틸렌 또는 나일론 백. 백은 정전기 발생 경향이 낮아야 하며, 활택제나 가소제와 같은 가공 첨가제가 없어야 합니다. 백 자체에 대한 인증서를 제공하십시오.
- 내부 봉투 밀봉: 세척 및 건조된 부품을 내부 백에 넣으십시오. 필요한 경우, 완충 작용을 하는 불활성 기체를 만들기 위해 여과된 질소로 백을 퍼지하십시오. 열압착 밀봉기를 사용하여 백을 밀봉하여 견고하고 연속적인 밀봉을 만드십시오. 테이프, 스테이플러, 지퍼는 사용할 수 없습니다.
- 단계별 소독: 다음 포장 단계로 넘어가기 전에 밀봉된 내부 봉투의 외부를 멸균 처리된 보풀이 생기지 않는 폴리에스터 물티슈를 사용하여 이소프로필 알코올(IPA)과 증류수를 70/30 비율로 혼합한 용액으로 꼼꼼하게 닦아내야 합니다.
- 외부 봉투 밀봉: 낮은 등급의 환경(예: ISO 7)에서는 닦아낸 내부 백을 외부 백에 넣고 동일한 방식으로 밀봉합니다. 이렇게 하면 가운 착용실이나 자재 에어록에서 외부 백을 제거하고 깨끗한 내부 백을 메인 클린룸으로 반입할 수 있는 시스템이 구축됩니다.
Expert tip: 경우에 따라 내부 포장이 밀폐되어 있는 경우, 항공 운송 시 팽창 효과를 고려해야 합니다 (충분히 튼튼한 밀봉 또는 골판지 상자로 감싸거나 팽창 공간을 확보해야 합니다).
4. 포장에 있어 엄격한 '종이, 판지 사용 금지' 규칙 적용

종이나 판지는 부품 또는 해당 부품의 1차/2차 클린룸 백에 직접 닿아서는 안 됩니다.
- 라벨: 가방에 직접 부착하는 모든 식별 라벨은 아크릴 접착제가 있는 합성 소재(예: 폴리에스터)로 제작되어야 하며, 번짐이 없는 잉크로 인쇄되어야 합니다. 라벨은 외부 가방의 바깥쪽에 부착해야 합니다. 내부에 '여행자' 태그가 필요한 경우, 클린룸 등급의 합성 용지(예: 타이벡)에 인쇄해야 합니다.
- 선적 서류 비치: 적합성 인증서, 분석 보고서, 포장 명세서 등 모든 서류는 별도의 파우치에 넣어 제품에 부착해야 합니다. 외부 최종 밀봉 배송 상자에 담겨서 발송되거나 전자적으로 전송됩니다. 어떠한 경우에도 배송 상자 안에 종이를 넣어서는 안 됩니다.
5. 최종 선적 컨테이너 준비
최종 배송 용기(일반적으로 골판지 상자)는 심각한 오염원이 될 수 있으므로 깨끗하게 포장된 제품을 격리할 수 있도록 준비해야 합니다.
- 박스 내부 안감: 배송 상자 안쪽에는 먼저 새롭고 다용도로 사용할 수 있는 폴리에틸렌 라이너 백을 깔아야 합니다.
- 던니지: 이중 포장된 제품을 박스 라이너 안에 넣을 때는 클린룸 환경에 적합한 완충재(예: 새 폴리에틸렌 폼 또는 에어캡)만 사용하십시오. 포장용 스티로폼 알갱이, 잘게 찢은 종이 또는 크라프트지의 사용은 엄격히 금지됩니다.
- 밀봉: 이중 포장된 제품을 안감이 있는 상자 안에 넣은 후, 안감을 접어 덮고 일반 포장 테이프로 상자를 밀봉합니다. 이렇게 하면 상자를 열었을 때 받는 사람이 가장 먼저 만지는 것이 깨끗한 안감이고, 상자 먼지가 묻지 않습니다.
아웃트로

견고한 오염 제어 전략은 단순한 문서가 아니라 모든 클린룸 운영을 총괄하는 능동적인 핵심 요소입니다. 이는 품질 위험 관리 원칙을 자재 이송 및 무균 기술부터 검증된 소독 프로토콜에 이르기까지 모범 사례를 구체적이고 일상적으로 실행하는 데 반영합니다.
궁극적으로 전략의 성공 여부는 그 범위가 아니라 검증 가능한 결과, 즉 통제 상태를 확인하고 시스템의 지속적인 발전을 뒷받침하는 일관된 환경 모니터링 데이터로 측정됩니다.
자주 묻는 질문
How does a Contamination Control Strategy (CCS) differ from individual cleaning and gowning SOPs?
A CCS is the overarching strategic document that integrates all control elements based on risk. SOPs are the detailed, tactical instructions for executing specific tasks defined within that strategy.
What is the correct technique for wiping items for transfer into a Grade A/B area, and why is contact time critical?
The correct technique involves using sterile, low-lint wipes saturated with a validated disinfectant in overlapping, unidirectional strokes. Contact time is critical because it is the validated duration required for the disinfectant to achieve its sporicidal or bactericidal action.
What are the most critical data inputs for the periodic review and update of a CCS?
The most critical inputs are environmental and personnel monitoring trend data, process deviations, media fill results, and CAPA effectiveness data. These inputs verify control and highlight areas needing reassessment.
What is the best practice for integrating supplier and raw material controls into the facility’s CCS?
The CCS must reference the supplier qualification program, which mandates audits, quality agreements, and microbial specifications for incoming materials. This extends contamination control to the entire supply chain.
Why are unidirectional, overlapping strokes mandated for cleaning and disinfecting surfaces in a cleanroom?
This technique ensures complete surface coverage and prevents the re-contamination of areas that have already been cleaned. It physically lifts and removes contaminants rather than just spreading them around the surface.
What are the key validation requirements for cleaning and disinfection agents cited in a CCS?
Validation requires efficacy studies (coupon studies) using in-house microbial isolates on representative manufacturing surfaces. These studies must prove the effectiveness of the agents at their specified contact times.
What are the primary differences in a CCS for an aseptic process versus a terminally sterilized product?
An aseptic process CCS focuses intensely on the exclusion of all microorganisms, as there is no final kill step. A CCS for terminal sterilization focuses on controlling the pre-sterilization bioburden to a validated limit.
What is the correct sequence for de-gowning, and why is it as critical as the gowning process itself?
De-gowning proceeds from the “dirtiest” items to the “cleanest,” typically starting with gloves and moving inward to the coverall, to contain contaminants on the gown. This prevents the operator from contaminating their scrubs or skin and dispersing particles from the used gown into the changing room.
Related Readings
- Aseptic Process Simulation (Media Fill) design and interpretation: it involves designing worst-case simulations to qualify a process’s aseptic integrity. Interpreting the results is critical for validating the process and qualifying operators.
- 미생물 식별 전략 및 데이터 동향: this deals with identifying environmental isolates to the species level to understand the facility’s microflora. Trending this data is essential for detecting shifts that indicate a loss of control.
- Design and control of RABS and isolator technology: it explores the validation and operation of advanced barrier systems. It includes integrity testing and glove management to ensure a superior aseptic environment.
- WFI and Pure Steam System validation and biofilm control: it shall cover the design and routine monitoring of high-purity water systems. A primary focus is on system validation and ongoing strategies to prevent and control biofilm formation.
- Container Closure Integrity Testing (CCIT) methodologies: the technologies used to ensure the final product seal is integral, guaranteeing sterility until use. It includes methods like vacuum decay and high voltage leak detection.
- Supplier and raw material microbial control programs: it shall extend contamination control to the supply chain through supplier audits and quality agreements. It involves setting and verifying microbial specifications for all incoming materials.
- Sterilization validation for components and equipment: validating the lethality and repeatability of sterilization cycles (e.g., autoclave, dry heat) for all items entering the aseptic processing area.
- Management of disinfectant and cleaning agent residues: it addresses the potential for chemical residues to inhibit disinfectant efficacy or become a product contaminant. It covers strategies for residue detection, removal, and rotation of cleaning agents.
External Links on Contamination Control Strategy (CCS)
국제 표준
- ISO 14644-1: 2015 Cleanrooms and controlled environments – Part 1: Classification of air cleanliness
- ISO 14644-2: 2015 Cleanrooms and controlled environments – Part 2: Monitoring to provide evidence of cleanroom performance related to air cleanliness by airborne particulate contamination
- ISO 14644-3: 2019 Cleanrooms and controlled environments – Part 3: Test methods
(링크 위에 마우스를 올려놓으면 콘텐츠에 대한 설명을 볼 수 있습니다.)
사용된 용어집
Bioburden: the presence of viable microorganisms on a surface or in a substance, typically measured to assess contamination levels in pharmaceuticals, medical devices, and other sterile products. It is crucial for determining sterilization efficacy and product safety.
Building Management System (BMS): a centralized control system that monitors and manages a building's mechanical and electrical equipment, including heating, ventilation, air conditioning, lighting, security, and fire safety systems, to enhance operational efficiency, comfort, and energy management.
Carbon Capture & Sequestration (CCS): a process that captures carbon dioxide emissions from sources like power plants and industrial processes, transporting it for storage underground or utilizing it in various applications, thereby reducing greenhouse gas concentrations in the atmosphere.
Certificate of Analysis (CoA): a document issued by a manufacturer or testing laboratory that confirms a product's specifications, quality, and compliance with regulatory standards, detailing test results and methods used for analysis.
Certificate of Conformance (CoC): 제조업체 또는 공급업체가 제품이 특정 기준, 규정 또는 계약 요건을 충족함을 확인하는 문서로, 품질 보증 및 규정 준수 검증 과정에서 자주 사용됩니다.
Code of Federal Regulations (CFR): 미국 연방 기관에서 발행한 일반적이고 영구적인 규칙들을 주제별로 50개 편으로 정리한 것으로, 연방 규정에 대한 공식적인 법적 자료 역할을 합니다.
Colony Forming Units (CFU): a measurement used to estimate the number of viable microorganisms in a sample, indicating the number of cells capable of forming colonies under specific growth conditions.
Computational Fluid Dynamics (CFD): a numerical method used to analyze fluid flow, heat transfer, and related phenomena by solving the governing equations of fluid motion and thermodynamics through discretization techniques, enabling simulation and visualization of complex fluid behavior in various engineering applications.
Contamination Control Strategy (CCS): a systematic approach to prevent, detect, and mitigate contamination in controlled environments, ensuring product quality and safety through defined procedures, monitoring, and risk management practices.
Corrective Action and Preventative Action (CAPA): a systematic approach to identifying, investigating, and addressing nonconformities and potential issues to prevent recurrence and ensure compliance with regulatory standards in quality management systems.
Corrective and Preventive Action (CAPA): 규제 기준 준수 및 프로세스와 제품의 지속적인 개선을 보장하기 위해 부적합 사항을 식별, 조사 및 해결하는 체계적인 접근 방식입니다.
Critical Control Points (CCP): specific stages in a process where control can be applied to prevent, eliminate, or reduce food safety hazards to acceptable levels. Identifying these points is essential for effective hazard analysis and critical control management in food production systems.
current Good Manufacturing Practice (cGMP): a system ensuring that products are consistently produced and controlled according to quality standards, encompassing regulations and guidelines for manufacturing processes, facilities, equipment, and personnel to ensure safety, quality, and efficacy in pharmaceuticals, food, and other regulated industries.
Electronics Manufacturing Services (EMS): 아웃소싱 서비스는 원장비 제조업체(OEM)를 위해 전자 부품 및 시스템의 설계, 생산, 조립 및 테스트를 제공하여 비용 효율적이고 효율적인 제조 공정을 가능하게 하는 동시에 고객이 핵심 사업 활동에 집중할 수 있도록 지원합니다.
Environmental Monitoring System (EMS): 대기, 수질, 토양 등 환경조건에 대한 데이터를 체계적으로 수집, 분석, 보고하여 법규 준수 여부를 평가하고, 시간에 따른 변화를 추적하며, 환경 관리 및 보호를 위한 의사결정을 지원하도록 설계된 시스템입니다.
Failure Mode and Effects Analysis (FMEA): a systematic method for evaluating potential failure modes within a system, process, or product, assessing their effects on performance, and prioritizing risks to improve reliability and safety through corrective actions.
Food and Drug Administration (FDA): a federal agency of the United States Department of Health and Human Services responsible for regulating food safety, pharmaceuticals, medical devices, cosmetics, and tobacco products to ensure public health and safety through scientific evaluation and enforcement of compliance standards.
Good Manufacturing Practice (GMP): a system ensuring products are consistently produced and controlled according to quality standards, minimizing risks involved in pharmaceutical production and related industries. It encompasses guidelines for manufacturing processes, facility conditions, personnel qualifications, and documentation practices to ensure product safety and efficacy.
Hazard Analysis and Critical Control Points (HACCP): a systematic approach to food safety that identifies, evaluates, and controls hazards at critical points in the production process to prevent foodborne illnesses and ensure product safety.
Heating Ventilation and Air Conditioning (HVAC): a system designed to regulate indoor climate by controlling temperature, humidity, and air quality through heating, cooling, and ventilation processes. It includes components such as furnaces, air conditioners, ductwork, and thermostats for efficient environmental management.
Heating, Ventilation, and Air Conditioning (HVAC): 난방, 냉방 및 환기 과정을 통해 온도, 습도 및 공기 질을 제어하여 실내 환경을 조절하도록 설계된 시스템으로, 주거, 상업 및 산업 환경에서 쾌적함과 안전을 보장합니다.
International Organization for Standardization (ISO): a non-governmental international body that develops and publishes standards to ensure quality, safety, efficiency, and interoperability across various industries and sectors, facilitating global trade and cooperation. Established in 1947, it comprises national standardization organizations from member countries.
Key Performance Indicator (KPI): a measurable value that demonstrates how effectively an organization is achieving key business objectives, often used to evaluate success at reaching targets.
Material Airlock (MAL): a sealed entryway designed to control the transfer of materials between different environments, preventing contamination and maintaining specific atmospheric conditions. It typically consists of two or more interlocking doors that ensure isolation during the transfer process.
Performance Qualification (PQ): a process that verifies a system or equipment operates according to specified requirements under real-world conditions, ensuring it consistently performs its intended function within predetermined limits.
Personnel Airlock (PAL): a sealed entryway designed to allow personnel to transition between different pressure environments while minimizing contamination and maintaining safety, typically used in space stations, laboratories, or cleanrooms. It features interlocking doors that prevent simultaneous opening.
Standard Operating Procedure (SOP): a set of step-by-step instructions created to help workers carry out routine operations consistently and efficiently, ensuring compliance with regulations and quality standards.
Statistical Process Control (SPC): a method of quality control that employs statistical techniques to monitor and control a process, ensuring it operates at its full potential by identifying variations and maintaining consistent output within specified limits.
Total Organic Carbon (TOC): a measure of the amount of carbon found in organic compounds within a sample, often used to assess water quality and environmental health. It quantifies the concentration of carbon derived from organic matter, excluding inorganic carbon sources.
Unique Selling Point (USP): a distinctive feature or benefit that sets a product or service apart from competitors, aimed at attracting customers by addressing specific needs or preferences.
Volatile Organic Compound (VOC): organic chemicals that have a high vapor pressure at room temperature, leading to significant evaporation and potential air pollution. They are commonly found in paints, solvents, and fuels, contributing to smog formation and adverse health effects.