A Contamination Control Strategy (CCS) is a comprehensive, documented framework that establishes the formal approach for managing risks to product quality from all forms of contamination. It is a living document that goes far beyond facility design, integrating disparate elements such as personnel training, material transfer protocols, process flows, environmental monitoring, and utility qualification into a single, cohesive plan. The CCS is not merely a summary of existing procedures; it is the strategic justification, based on risk management, that defines why specific controls are implemented, how they are monitored, and what actions are taken when they deviate from their intended state of control.
The cleanroom is one of the physical components, but it is fundamentally governed by the CCS. Its classification, operational parameters, and monitoring limits are direct outputs of the risk assessments detailed within the strategy. The CCS provides the rationale for the cleanroom’s existence and dictates its performance requirements to mitigate identified contamination risks.
Key Takeaways
- CCS should be built upon the principles of Quality Risk Management (QRM)
- Holistic, facility-wide implementation strategy.
- Define and monitor critical control points.
- Personnel are a primary contamination source.
- Proactive and continuous environmental monitoring.
- Implement strict material and waste controls.
- Process design to minimize exposure
- Ensure regulatory compliance (e.g., Annex 1).
What is a Contamination Control Strategy (CCS)?
A Contamination Control Strategy (CCS) is a formal, comprehensive, and living document that defines a manufacturer’s entire approach to minimizing contamination in their products.
Mandated by the revised EU GMP Annex 1, its core purpose is to move away from a fragmented collection of separate procedures (for cleaning, gowning, monitoring, etc.) towards a single, holistic strategy that demonstrates a deep understanding of all potential contamination risks and the scientific rationale for the controls put in place to mitigate them.
The philosophy behind the CCS is that sterility and product quality are not achieved by any single action, but by the cumulative effect of a series of interconnected controls. The CCS is the master document that identifies every potential source of contamination—particulate, microbial, endotoxin/pyrogen, and chemical—and describes how the combination of facility design, equipment, procedures, and monitoring programs work together to protect the product throughout its lifecycle.
Key Elements of a Contamination Control Strategy
1. Facility and Equipment Design
This element is the physical foundation of the Contamination Control Strategy. It’s not enough to have a cleanroom; you must justify why it was designed the way it was, based on risk.
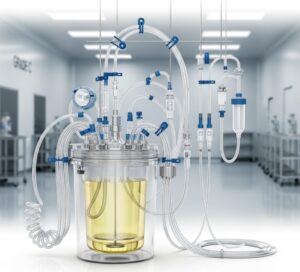
Rationale for the design: the CCS must detail the logic behind the facility’s layout, including unidirectional flows for personnel, materials, equipment, and waste to prevent mix-ups and backtracking from “dirty” to “clean” areas. It must scientifically justify the pressure cascades between rooms (e.g., the Grade B cleanroom is maintained at a significant positive pressure relative to the surrounding Grade C support area), providing data from the Building Management System (BMS) to prove these differentials are continuously maintained. The design of material and personnel airlocks, including their interlock mechanisms and purge times, must be rationalized as critical control points.
Equipment design: the Contamination Control Strategy justifies the selection of process equipment based on its ability to mitigate contamination. This includes specifying sanitary design features like crevice-free surfaces, 316L stainless steel, and Tri-Clamp fittings to prevent microbial colonization and facilitate effective cleaning. Crucially, it details the rationale for using advanced aseptic technologies. For example, it would justify the use of a Restricted Access Barrier System (RABS) or a fully contained isolator by referencing a risk assessment (FMEA) that shows these technologies significantly reduce the risk of operator-borne contamination compared to a conventional open-process cleanroom.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1: “Manufacture of Sterile Medicinal Products” (August 2022), specifically Section 4, “Premises.”
- FDA: Code of Federal Regulations, Title 21, Part 211.42, “Design and construction features” and 211.46, “Ventilation, air filtration, air heating and cooling.”
Tip: use Computational Fluid Dynamics (CFD) modeling during the design phase of a new cleanroom or the installation of new equipment. Supplement this with physical smoke studies during qualification. The visual evidence from these studies provides irrefutable, science-based proof that your airflow patterns are unidirectional, effectively protecting the critical zone from contamination, which is far more powerful during an audit than simply presenting air velocity and pressure differential data.
2. Personnel
This element recognizes that humans are the primary source of microbial and particulate contamination in a cleanroom.
Gowning: the Contamination Control Strategy details the entire gowning system, not just the procedure. This includes the material science behind the chosen garments (e.g., non-shedding, fluid-resistant), the validation of their sterilization cycle, and the results of gowning qualification studies for each operator. These studies must provide objective data (e.g., contact plates from gloves and sleeves) to prove that an individual can gown without compromising the sterility of the garment.
Training and aseptic technique: the strategy goes beyond simple procedural training. It describes a formal qualification program where operators must demonstrate proficiency in aseptic manipulations, often through successful participation in media fills (aseptic process simulations). The CCS emphasizes the “why” behind the rules, ensuring personnel understand the microbiological principles of contamination control, such as the importance of slow, deliberate movements to avoid generating air turbulence.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 7, “Personnel.”
- FDA: Code of Federal Regulations, Title 21, Part 211.28, “Personnel responsibilities” and 211.113, “Control of microbiological contamination.”
Tip: implement a “human factors” study for your most critical aseptic interventions. Instead of just writing a procedure, observe operators performing the task and analyze the ergonomics and cognitive load. A procedure that is difficult, awkward, or confusing is a contamination risk. Simplifying the process based on human factors engineering is a proactive control that is far more effective than simply retraining personnel on a flawed procedure.
3. Utilities
Utilities that contact the product or product-contact surfaces are a direct potential pathway for contamination.
Control of critical utilities: the Contamination Control Strategy provides a comprehensive overview of the design, validation, and ongoing monitoring of critical utility systems. For a Water-for-Injection (WFI) system, it would detail the continuous hot-loop circulation (>80°C) to prevent biofilm formation, the validation of the multi-stage purification process, and the routine monitoring program for bioburden, endotoxin, Total Organic Carbon (TOC), and conductivity. Similarly, it would describe the filtration (e.g., sterile 0.22 µm filters) and qualification of process gases like compressed air or nitrogen that come into direct product contact.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 6, “Utilities.”
- FDA: Code of Federal Regulations, Title 21, Part 211.63, “Equipment design, size, and location,” supported by the principles in the “Guidance for Industry: Sterile Drug Products Produced by Aseptic Processing — Current Good Manufacturing Practice.”
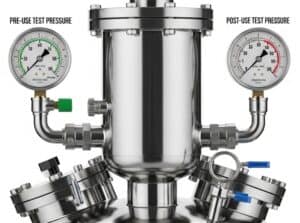
Application example: the CCS for a biologics facility details the risk assessment for the process air system. It identifies the point-of-use sterile filters on the bioreactor air inlet as Critical Control Points. The strategy can mandate an integrity test of these filters not only before the batch (pre-use) but also after the batch is harvested (post-use, pre-sterilization) to prove the filter remained integral throughout the entire cell culture process, thus ensuring the batch was protected.
Tip: use online Total Organic Carbon (TOC) monitoring in your WFI loop not just as a quality attribute test, but as a leading indicator for system health. Establish a tight internal alert limit, well below the official action limit. A slow, upward trend in TOC levels is often the very first sign of developing biofilm in the system, allowing maintenance to intervene with sanitization cycles long before a significant microbial or endotoxin event occurs.
4. Raw Materials and Components
Contamination control begins with the materials that enter the facility.
Bioburden and endotoxin control: the CCS outlines a proactive strategy for managing the microbial quality of all incoming materials. This involves setting specifications for bioburden and endotoxin levels on raw materials, primary packaging (vials, stoppers), and product-contact components. This data is critical, as it is used to validate the robustness of downstream removal or sterilization steps (e.g., proving an autoclave cycle can achieve the required Sterility Assurance Level based on the maximum potential starting bioburden).
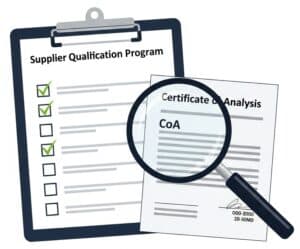
Supplier management: this section details the risk-based supplier qualification program. It describes how suppliers of critical components are audited and qualified, and the role of formal Quality Agreements in defining quality responsibilities. It also specifies the program for periodically verifying a supplier’s Certificate of Analysis (CoA) through in-house testing to ensure ongoing consistency and control.
Regulatory References:
- EU: EudraLex – Volume 4 – Part I, Chapter 5, “Production” and Chapter 7, “Outsourced Activities.”
- FDA: Code of Federal Regulations, Title 21, Part 211.84, “Testing and approval or rejection of components, drug product containers, and closures.”
Tip: implement a risk-based program for your incoming component testing that goes beyond the standard pharmacopeia. For stoppers used in sensitive biologic drug products, your CCS should include a testing regime for extractables and leachables under worst-case conditions to ensure that no compounds that could degrade the drug molecule are introduced from the component itself.
Bioburden Details
The bioburden refers to the population of viable microorganisms present on a surface, in a liquid, or within a raw material before it undergoes a sterilization or disinfection process.
This microbial load, measured in Colony Forming Units (CFU), is a key indicator of the cleanliness of the manufacturing environment and a crucial factor in ensuring product safety and efficacy.
The primary goal in these industries is to maintain a low bioburden to minimize the challenge to the final sterilization process and to prevent product contamination. High bioburden levels can compromise the sterility of a product, potentially leading to infections in patients or defects in sensitive electronic components.
Bioburden in the pharmaceutical industry The pharmaceutical industry operates under stringent regulatory requirements to control microbial contamination. The United States Pharmacopeia (USP) provides specific guidelines for bioburden limits in both sterile and non-sterile drug products. Non-sterile products: for non-sterile pharmaceuticals, the acceptable bioburden levels vary depending on the product’s intended use and route of administration. USP chapter <61> outlines the microbial enumeration tests, while chapter <1111> provides the acceptance criteria. These criteria are designed to ensure that the product is safe for its intended use and that the microorganisms present will not proliferate to harmful levels. Sterile products: for sterile drug products, the bioburden must be meticulously controlled before the final sterilization process. According to USP chapter <62>, sterile products should have a total aerobic microbial count (TAMC) and total yeast and mold count (TYMC) of no more than 10 CFU per 100 mL. The European Medicines Agency (EMA) also recommends a pre-sterilization bioburden limit of less than 100 CFU per 100 mL. | Bioburden in the medical device industry Similar to pharmaceuticals, the medical device industry places a strong emphasis on controlling bioburden to ensure the effectiveness of sterilization and the safety of the device. The international standard ISO 11737-1 provides guidance on the determination of the population of microorganisms on medical devices. The acceptable bioburden for a medical device can vary depending on its classification, intended use, and the materials it is made from. While there are no universal, mandated limits for all devices, a typical bioburden level for a medical device is between 0 and 150 CFUs. For some devices, a lower limit of less than 100 CFU may be targeted to ensure a higher sterility assurance level. The size and complexity of the device can also influence its bioburden, with larger and more intricate devices potentially having higher microbial loads. |
5. Cleaning and Disinfection
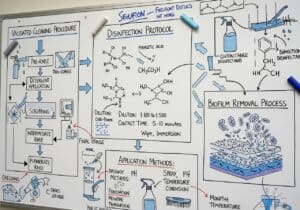
This element describes the chemical and physical removal of contaminants from surfaces (cleaning) and living organisms removal (disinfection).
Validated procedures: the CCS provides the scientific justification for the entire disinfection program. This includes the rationale for selecting specific disinfectants (e.g., a broad-spectrum bactericide) and sporicides (e.g., a peracetic acid-based agent), supported by coupon studies that prove their efficacy against the facility’s common microbial isolates. It details the validated parameters for their use, including dilution, contact time, and application method.
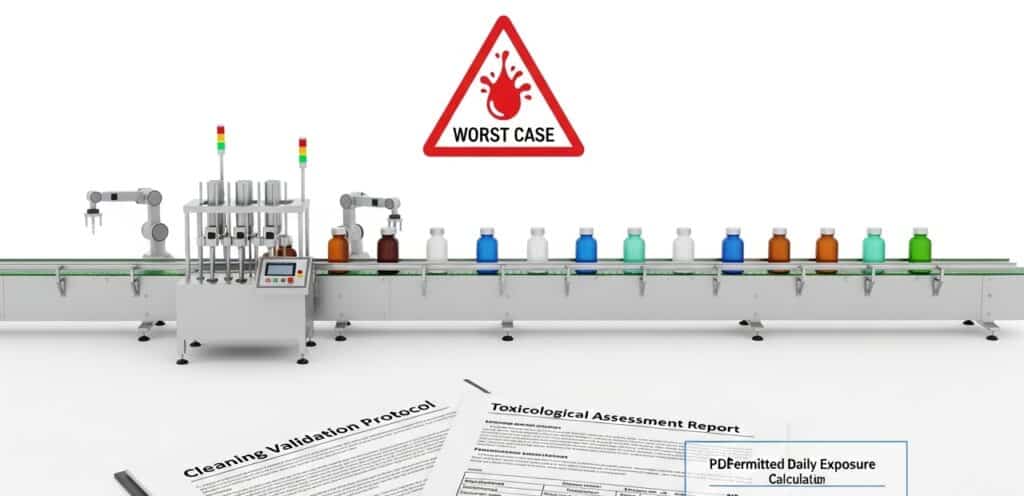
Efficacy and rotation: the strategy must include a disinfectant rotation program to prevent the development of resistant microbial strains. The CCS explains the frequency and logic of this rotation. It also describes the validation of cleaning procedures for product-contact equipment, proving they can effectively remove both chemical residues of the previous product and any microbial contamination to pre-defined, health-based limits.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 5, “Cleaning and disinfection.”
- FDA: Code of Federal Regulations, Title 21, Part 211.67, “Equipment cleaning and maintenance.”
Tip: your disinfectant efficacy validation should not be a one-time event. Your Contamination Control Strategy should mandate an ongoing “in-situ” efficacy program. Periodically, take environmental isolates (the actual bacteria and molds growing in your facility) and re-challenge your validated disinfectants in the lab. This verifies that your program remains effective against the current, relevant microflora, which can change over time.
6. Process Risk Management
This is the intellectual core of the CCS, where all potential hazards are formally identified and controlled.
Risk identification: the CCS must contain or reference a formal process risk assessment, typically an FMEA or HACCP. This assessment systematically breaks down the entire manufacturing process, step-by-step, to identify every potential contamination risk (e.g., aseptic connection, operator intervention, material transfer).
Critical Control Points: based on the risk assessment, the CCS identifies the Critical Control Points (CCPs)—the specific steps where control is essential to prevent or eliminate a risk. For each CCP, the CCS defines the specific control measure (e.g., use of a single-use sterile connector, continuous particle monitoring during filling) and the scientific rationale for why that control is effective.
Quality risk management always comes back to the risk benefit ratio of each control measure: a stricter cleanroom practice is justified only when its protection outweighs its cost. Teams that document the risk to benefit ratio of every control can defend their choices during an audit.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 2, “Pharmaceutical Quality System (PQS),” which explicitly requires the application of Quality Risk Management (ICH Q9).
- FDA: The principles are embedded throughout the Aseptic Processing Guidance, which is built on a foundation of risk management. The FDA has formally adopted the ICH Q9 guideline.
Tip: make your process risk assessments “living documents.” The CCS should mandate a formal review of the FMEA for a given process after a set period (e.g., one year) or after a set number of batches. During this review, use actual deviation and batch failure data to re-score the “Probability” and “Detection” elements of the FMEA. This transforms the FMEA from a theoretical exercise into a dynamic, data-driven tool that accurately reflects the real-world performance of your process.
7. Environmental and Process Monitoring
This element describes the systems used to verify that the facility and process remain in a state of control.
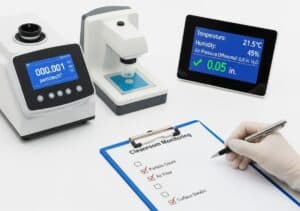
The EM program: the CCS provides a detailed rationale for the Environmental Monitoring (EM) program. It justifies every sampling location based on the process risk assessment (e.g., locations where sterile product is exposed, areas of frequent operator activity). It defines the methods (e.g., active air sampling, settle plates, contact plates), frequencies, and alert/action limits for both non-viable particulates and viable microbial counts.
Process monitoring: this section describes the use of modern monitoring technologies. This includes continuous, real-time particle monitoring in the Grade A critical zone, with alarms linked to the Batch Record. It also describes the trending of pressure differentials and other critical parameters. The Contamination Control Strategy must also define the program for identifying microbial isolates recovered from the EM program to the species level, which is crucial for tracking the facility’s microbial flora and investigating deviations.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 9, “Viable and non-viable environmental and process monitoring.”
- FDA: Code of Federal Regulations, Title 21, Part 211.113(b), which requires “appropriate written procedures… designed to prevent microbiological contamination.” The Aseptic Processing Guidance provides detailed expectations.
Tip: use the microbial identification data from your EM program to create a “microbial map” of your facility. Track not just the counts, but the species and where they are found. This allows you to identify resident “house” organisms and track their movement. If a media fill or sterility test failure occurs, you can compare the contaminating organism to your facility map, which can dramatically accelerate the root cause investigation by pointing to the likely source area.
8. Continuous Improvement
The CCS is not a static document; it must evolve and is highly related to the “c” of the cGMP.
Data trending: the CCS mandates a formal system for the routine trending of all data related to contamination control, including EM results, personnel monitoring, utility monitoring, and batch-related deviations. Statistical Process Control (SPC) should be used to identify negative trends, even if the results are still within specification.
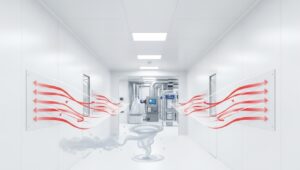
Feedback loop: this is the most critical part of the continuous improvement cycle. The Contamination Control Strategy must be a formal agenda item in the site’s Quality Management Review. The trend data must be reviewed by senior management, and the output must be a set of documented actions for improvement. If a new microbial strain is frequently recovered, for example, the CCS must be updated to reflect a potential change in the disinfection program or an investigation into its source.
Regulatory References:
- EU: EudraLex – Volume 4 – Annex 1 (August 2022), Section 2.5, which states the CCS “should be actively updated and should drive continuous improvement.” This links directly to ICH Q10, “Pharmaceutical Quality System.”
- FDA: The principles of continuous improvement are central to the PQS model described in ICH Q10, which the FDA has adopted.
Tip: integrate the CCS review directly with the site’s Quality Management Review (QMR) and use a “Quality Risk Management” dashboard. Present KPIs derived from the CCS (e.g., EM excursion rates, media fill pass rates, contamination-related deviation trends) directly to senior leadership. This ensures that the health of your contamination control program has high-level visibility, which is critical for securing the resources and management commitment needed to drive meaningful continuous improvement.
Cleanrooms
Cleanrooms are an important part of the Contamination Control Strategy (CCS), together with logistics procedure and suppliers management.
Disclaimer: the following chapters are intended for design, quality and process users to better apply cleanroom principles. Despite being very detailed, cleanroom management implies many factors and specificity due to the variety of products and requirements; dedicated professionals and suppliers are required for this.
Cleanroom Classifications
This table compares the most common cleanroom classes under the international ISO 14644-1 standard and the European Union’s EudraLex Volume 4, Annex 1 (GMP) standard.
ISO Class 5 EU GMP Grade A | ISO Class 6 | ISO Class 7 EU GMP Grade B | ISO Class 8 EU GMP Grade C / D | |
| Airflow Characteristics | Unidirectional Flow (UDF) Also known as Laminar Flow: air moves in a single direction at a consistent velocity (typically 0.36–0.54 m/s). | Non-Unidirectional Flow Also known as Turbulent Flow: contamination is controlled by dilution. | ||
| Primary Function | Critical Zone Protection Designed to actively protect exposed sterile products and surfaces from contamination. | High-Quality Process Control Used for processes requiring very low particle counts but not necessarily full aseptic conditions. | Aseptic Process Support Serves as the immediate background environment for a Grade A zone. | General Clean Support Serves as a background for less critical activities or as an entry/gowning area for higher classes. |
Max Particle Concentration (In Operation) | ISO 5:
Grade A:
|
| ISO 7:
Grade B:
| ISO 8 / Grade C:
Grade D: not defined in operation. |
Max Particle Concentration (At Rest) | ISO 5 / Grade A:
|
| ISO 7 / Grade B:
| ISO 8 / Grade C:
Grade D:
|
Microbial Limits (In Operation) | Air Sample: <1 CFU/m³ Settle Plate (4 hrs): <1 CFU Contact Plate: <1 CFU | Not formally defined by ISO, application-dependent. | Air Sample: 10 CFU/m³ Settle Plate (4 hrs): 5 CFU Contact Plate: 5 CFU | Grade C: Air: 100, Settle: 50, Contact: 25 Grade D: Air: 200, Settle: 100, Contact: 50 |
| Typical Air Changes / Velocity | Velocity: 0.36 – 0.54 m/s (90 ft/min ± 20%) (ACH is not the primary metric for UDF) | ACH: 150 – 240 | ACH: 60 – 90 | ACH: 20 – 45 |
| Required Dressing | Full Sterile Gowning: Sterile, individually wrapped hood, goggles, mask, coverall, and overboots. Two pairs of sterile gloves; inner pair tucked under cuff, outer pair pulled over. Outer gloves regularly sanitized. | Full Gowning: Clean-processed (not necessarily sterile) coverall, hood, boots, gloves, and mask. | Full Sterile Gowning: Identical to Grade A. All items must be sterile. | Grade C: Dedicated scrubs or coverall, dedicated shoes/overshoes, hairnet, beard cover, mask, gloves. Grade D: Clean plant uniform, dedicated shoes or shoe covers, hairnet, beard cover. |
| Must-Have Hardware | • Filtering: Full coverage of terminal HEPA/ULPA filters to achieve unidirectional flow. • Surfaces: Pharmaceutical-grade, non-porous, non-shedding, smooth surfaces (sandwich panels, poured epoxy/welded vinyl floors). • Construction: All joints fully coved and sealed. No ledges. | • Filtering: Terminal HEPA filters in the ceiling. • Surfaces: Hard, non-porous, smooth surfaces. • Construction: Coved flooring. | • Filtering: Terminal HEPA filters in the ceiling. • Surfaces: Hard, non-porous, non-shedding, smooth surfaces. • Construction: Interlocked airlocks for personnel/materials; visible differential pressure gauges. | • Filtering: HEPA filters (often in AHU or ducted). • Surfaces: Smooth, cleanable, non-porous floors and walls. • Construction: Defined gowning area/airlock; visible differential pressure gauges. |
| Nice-to-Have Hardware | • Integrated, real-time particle counters with alarms. • Automated fogging systems for bio-decontamination. • Integrated glove integrity testers for isolators. • Fully integrated wall/floor coving. | • Continuous particle monitoring system. • Interlocked pass-through chambers with HEPA filtration. • Chemical-resistant flooring. | • Real-time particle monitoring near critical areas. • Automated high-speed roll-up doors. • Wall panels with embedded utilities to reduce clutter. | • Networked particle counters. • Automated shoe cover applicators. • Wall-mounted stainless steel furniture. |
| Typical Applications | • Aseptic filling of vials • Sterile connections & transfers • Open processing of sterile materials • Cell therapy processing | • High-end medical device assembly • Non-sterile but low-bioburden pharmaceutical filling • High-quality optics assembly | • Background room for Grade A zones • Buffer and media preparation • Aseptic component assembly | • Grade C: Component washing, less critical aseptic steps, background for Grade B. • Grade D: Gowning rooms, less critical formulation, initial prep stages. |
| Governing Standards | US/International: ISO 14644-1:2015 EU: EudraLex Vol. 4, Annex 1 (2022) | US/International: ISO 14644-1:2015 | US/International: ISO 14644-1:2015 EU: EudraLex Vol. 4, Annex 1 (2022) | US/International: ISO 14644-1:2015 EU: EudraLex Vol. 4, Annex 1 (2022) |
| Typical Contaminants & Sizes | The standards measure total particle counts, not specific types. However, the primary contamination sources and their typical sizes are:
| |||
Contamination Comparison
To appreciate the stringency of cleanroom standards, it’s helpful to compare their particle count limits to those of everyday environments (these are typical values and of course can vary extremely):
| Office | Room at Home | Kitchen (During Cooking) | Normal Factory | |
| Typical Particle Count (per m³) | ~1,900,000 (≥0.5 µm) | >100,000,000 (≥0.3 µm) | Extremely high spikes, potentially >85,000,000,000 | Highly variable, often >35,000,000 |
| Common Particle Sizes | Micron-sized dust and fibers | Wide range, including allergens and sub-micron particles | Primarily ultrafine particles (<0.1 µm) and fine particles (PM2.5) | Process-dependent, from large dust to fine sub-micron particles |
| Primary Sources | Dust, clothing/carpet fibers, skin flakes, HVAC intake | Dust, pet dander, pollen, mold spores, human activity | Combustion, aerosolized oils and fats from cooking (frying, grilling) | Industrial processes (grinding, cutting), raw materials, combustion byproducts |
Conclusion: better work in the office than at home, especially if you have some allergies 😉
The ultimate cleanroom: the “Memory Facility” of the great Blade Runner 2049 movie, by Denis Villeneuve, based on Philip K. Dick’s book.
Cleanroom Best Design Practices
From pharmaceuticals to semiconductor manufacturing, the principles of cleanroom design are rooted in a multi-faceted approach for everything from airflow management to the selection of construction materials:
Strategic Layout and Workflow OptimizationA well-designed layout is crucial for minimizing contamination risks and improving operational efficiency. The design should enforce proper procedures and prevent cross-contamination by dictating the movement of people and materials. Zoning and adjacency: design the layout with clearly defined zones of increasing cleanliness, moving from external areas to gowning rooms, and finally into the main cleanroom. This creates a buffer and helps protect the most critical areas. Unidirectional flow: plan for a one-way flow of personnel, materials, and waste to prevent cross-contamination. This means establishing separate entry and exit points where feasible and ensuring that dirty materials do not cross paths with clean ones. | Meticulous HVAC System DesignThe Heating, Ventilation, and Air Conditioning (HVAC) system is the heart of a cleanroom, responsible for maintaining air cleanliness, temperature, humidity, and pressure. It can account for 50-80% of the facility’s energy use. Filtration levels: high-efficiency particulate air (HEPA) filters, which are 99.97% efficient at trapping particles of 0.3 microns, are standard. For the most stringent classifications (ISO 5 and below), ultra-low penetration air (ULPA) filters may be required to capture even smaller particles. Airflow patterns: the choice between unidirectional (laminar) and non-unidirectional (turbulent) airflow depends on the required ISO classification. Unidirectional flow, where air moves in a single pass, typically downwards, is used for stricter classes like ISO 5 to sweep particles away from critical processes, while turbulent flow is suitable for less critical environments (ISO 6 and above). |
Prioritize Contamination ControlEffective contamination control strategies are a top priority and must be integrated into the design from the outset. This involves a holistic approach that considers all potential sources of contamination, including personnel, materials, and equipment. Air showers: for highly sensitive cleanrooms, an air shower can be installed at the entrance to the gowning area. High-velocity jets of HEPA-filtered air blow off particulate contaminants from personnel before they enter the cleaner space. Strict gowning protocols: the design must support rigorous gowning procedures. This includes providing adequate space and facilities for personnel to don non-shedding cleanroom garments in the correct sequence. |  |
Judicious Selection of MaterialsThe materials used in construction must be non-shedding, non-porous, and easy to clean, while also being resistant to cleaning agents and disinfectants. Common material choices: stainless steel is often preferred for its durability, smooth surface, and resistance to corrosion and chemicals. Other common materials include epoxy-coated surfaces, fiberglass-reinforced plastic (FRP), and coated aluminum for wall panels. Flooring considerations: flooring must be seamless and durable. Vinyl and epoxy flooring are popular choices because they can be installed without joints, reducing the risk of particle accumulation, and can offer anti-static properties crucial for electronics manufacturing. | Seamless and Sealed SurfacesTo prevent the accumulation of particles and facilitate effective cleaning, all joints, seams, and penetrations in walls, floors, and ceilings must be properly sealed. Coved transitions: use coving where floors meet walls and where walls meet ceilings. This creates a curved, seamless transition that is easier to clean and prevents the buildup of contaminants in sharp corners. Flush integration: windows, light fixtures, and utility outlets should be designed to be flush with the wall or ceiling surfaces. This eliminates ledges and crevices where particles can settle and makes cleaning more effective. |
 | Controlled Environment ParametersPrecise control of temperature and humidity is critical not only for many sensitive processes but also for personnel comfort and contamination control. Humidity’s dual threat: high humidity (typically above 60%) can encourage microbial growth and cause corrosion on sensitive equipment. Conversely, low humidity (below 30%) can lead to the buildup of static electricity, which can damage electronic components and cause particles to cling to surfaces. Temperature and personnel: uncomfortable temperatures can cause personnel to sweat or shiver, increasing the shedding of particles and compromising the effectiveness of their cleanroom garments. A typical target is 21°C (70°F) with a small variance. (also see our “industry secret, that nobody is proud about” in 11. Gloved Hands Routine below and the relation with the room temperature control to limit sweating, no sharp edge or corner nor burr) |
Positive and Negative PressurizationMaintaining appropriate air pressure differentials between adjacent areas is crucial for preventing the ingress of contaminants. Air naturally flows from areas of higher pressure to lower pressure. Cascading pressure: a common strategy is a cascading pressure differential, where the cleanest room has the highest pressure. Each successive outer room or airlock has slightly lower pressure, ensuring that air always flows out of the cleanest spaces, preventing contaminants from entering. Bubble, sink, and cascade airlocks: different pressure schemes can be used depending on the goal. A “bubble” airlock has higher pressure than its surrounding rooms to protect the airlock’s contents. A “sink” airlock has lower pressure to contain contaminants within it. A “cascade” airlock has an intermediate pressure between a high-pressure cleanroom and a low-pressure corridor. | Modular and Flexible DesignIncorporating modular elements into the design allows for future expansion, relocation, and adaptation to changing process needs with minimal disruption, but must be carefully evaluated (see tip below). Reduced construction time: because components are prefabricated off-site, on-site assembly is significantly faster than traditional construction, reducing project timelines and downtime. Scalability and reconfiguration: modular cleanrooms can be more easily expanded or reconfigured if process requirements change. This adaptability makes them a cost-effective long-term investment. Tip: while we usually recommend wheels for equipment and benches for lean manufacturing by improving the modularity of production … this is to be minimized here due to the particules likely generated by the wheels or their brakes. |
Integrated and Cleanroom-Compatible LightingLighting must provide adequate illumination for detailed tasks without compromising the cleanroom’s integrity. Sealed and flush fixtures: light fixtures must be fully sealed to prevent them from collecting or shedding particles. They should be installed flush with the ceiling, eventually with a “teardrop” shape, to avoid disrupting the airflow pattern. |  |
Thoughtful Gowning Room DesignThe gowning room, or ante-room, serves as a critical buffer zone between the outside environment and the cleanroom. Its design is crucial for effective contamination control. Demarcation and flow: a well-designed gowning room has a clear line of demarcation, often including a “sit-over bench,” that separates the “dirty” side from the “clean” side. The layout should guide personnel through a logical, linear gowning sequence. Proper amenities: the room must be equipped with cleanroom-specific furniture for storing garments and facilitating the gowning process, such as lockers, dispensers for gloves and masks, and mirrors for self-inspection. | Minimized Furniture and EquipmentThis are the Lean Principles and the 5S, but applied to their maximum! Only essential furniture and equipment should be placed inside the cleanroom to reduce potential sources of contamination and surfaces that require cleaning. Cleanroom-grade materials: all items must be made from non-shedding, non-porous materials like stainless steel or polypropylene that are easily sanitized. Ergonomic and minimalist design: furniture should be designed to have minimal horizontal surfaces where particles can accumulate. It should also be ergonomically designed to support efficient and safe work practices. No stock, no spare, no paper instructions … |
 | Monitoring SystemsInstall a robust monitoring system to continuously track and record critical environmental parameters, ensuring ongoing compliance and allowing for immediate corrective action. Key parameters to monitor: these systems track airborne particle counts, air pressure differentials, temperature, and humidity in real-time. Control versus monitoring: it’s important to differentiate between a Building Management System (BMS), which actively controls the HVAC to maintain setpoints, and an Environmental Monitoring System (EMS), which independently records data for quality assurance and regulatory compliance, triggering alarms when parameters go out of specification. |
Dedicated Personnel Airlocks (PAL)A Personnel Airlock (PAL) is a specialized room that acts as the primary barrier for staff moving between an uncontrolled or lower-grade area and the cleanroom. Its purpose is to control contamination introduced by people. Interlocked doors: a critical feature of a PAL is an interlocking door system that prevents both the entry and exit doors from being open simultaneously. This maintains the pressure differential between the two areas and prevents a direct path for airborne contaminants to enter the cleaner space. Gowning and de-gowning procedures: the PAL is often synonymous with the gowning room, designed to facilitate the proper sequence of putting on and removing cleanroom garments. The design should physically guide the user through the process to minimize error. | In-and-Out for Goods and Materials (MAL)A Material Airlock (MAL) is a dedicated space designed for transferring materials, equipment, and finished products in and out of the cleanroom without compromising its integrity. Pass-throughs and larger airlocks: for smaller items, a wall-mounted pass-through box with interlocked doors is an efficient solution. For larger equipment or carts, a walk-in MAL is required, which functions similarly to a PAL but is designed for materials. Decontamination and wipe-down: the design must allow for materials to be decontaminated before they enter the cleanroom. This typically involves providing space and surfaces for operators to wipe down all items with appropriate sterile agents. Some advanced MALs may incorporate automated decontamination systems, such as vaporized hydrogen peroxide (VHP) foggers. |
Energy efficiency and sustainability focus
Cleanrooms are incredibly energy-intensive, so incorporating sustainable design principles can significantly reduce long-term operating costs and environmental impact. The main factor are thermal insulation and Variable Air Volume (VAV) systems: instead of running the HVAC system at full power continuously, a VAV system can adjust the airflow based on real-time conditions, such as particle counts or occupancy, reducing energy consumption during periods of low activity.
Tip: consider large windows or a glass tunnel if your industry requires frequent visitors, rather than letting them in (it will also show that you really care and are strict on procedures)
Bonus tip: as all materials are very specific, it may be less costly to build a cleanroom from scratch, including walls and ceiling, than to try desperately to upgrade an existing facility.
Cleanroom Best Human Practices
Excellent cleanroom design and state-of-the-art equipment as described above are only part of the equation for maintaining a sterile environment.
The single largest source of contamination in any cleanroom is the people working within it.
Therefore, rigorous adherence to specific human practices and behaviors is not just recommended—it is absolutely critical.
1. Gowning and De-Gowning SequenceThis is the first line of defense. Personnel must master the precise, step-by-step procedure for donning and removing cleanroom garments. “Cleanest to dirtiest” principle: gowning should follow a sequence that minimizes the contamination of sterile garments, typically starting from the head down (hood, mask, coveralls) and finishing with gloves, which are often donned last. No-touch technique: when donning sterile gloves, personnel must use a technique that ensures the sterile exterior of the glove is never touched by bare hands or any non-sterile surface. | 2. Deliberate and Controlled MovementAll movements inside the cleanroom must be slow, deliberate, and minimized to reduce the generation and dispersal of particles. The “cleanroom walk”: personnel should be trained to walk at a slow, steady pace without swinging their arms. Quick, jerky movements create turbulence that can stir up settled particles from surfaces. Minimize body motion: avoid unnecessary actions like stretching, scratching, or rapid head turning. All necessary movements should be performed efficiently and smoothly. |
3. Strict Personal HygieneContamination control begins before an operator even enters the gowning room. Prohibition of cosmetics and fragrances: makeup, hairspray, perfumes, and lotions can shed vast quantities of particles and volatile organic compounds (VOCs). These must be strictly forbidden. Pre-entry hand washing: hands and forearms must be thoroughly washed with a specified antimicrobial soap and dried with a non-shedding towel immediately before entering the gowning area. |  |
4. Proper “Cleanroom Posture”How a person stands and interacts with their workspace has a direct impact on contamination. Lean forward, not over: when working at a bench or process station, operators should lean from the hips and keep their head and torso back from the critical work area as much as possible. Never lean directly over the product or process. Respect the airflow: in a unidirectional (laminar) flow environment, operators must understand that their body is a significant source of contamination. They must always position themselves downstream of the critical product or process to ensure the clean, filtered air reaches the product first. | 5. “First Air” MentalityThis principle is paramount in unidirectional airflow cleanrooms. “First air” is the clean, filtered air that comes directly from the HEPA/ULPA filters before it passes over any object. Never block the airflow: operators must never place items, hands, or tools between the filter face and the critical product. This creates a “shadow” of turbulent, potentially contaminated air. Work in the cleanest zone: all critical manipulations should be performed in the direct path of the “first air,” as far upstream as possible. |
 | 6. “No-Touch” Policy for Exposed SkinEven with proper gowning, accidental exposure of skin can happen. Face and garment discipline: operators must be trained to never touch their face, adjust their mask or hood, or touch any part of their exposed skin while inside the cleanroom. If an adjustment is necessary, they must exit and re-gown. Glove integrity: gloves should be sanitized frequently with cleanroom-grade disinfectant (e.g., 70% isopropyl alcohol) and inspected regularly for tears or punctures. |
7. Cleanroom Communication MethodsSpeaking generates a significant amount of particle and droplet contamination. Minimize talking: verbal communication should be kept to an absolute minimum. Use non-verbal cues and systems: utilize hand signals, internal phone systems, or whiteboards for communication whenever possible. When speaking is necessary, it should be done facing away from the critical process. | 8. Material and Waste Flow ProtocolsThe way materials are handled is as important as how people move. “Triple-wrap” entry: materials are often brought into a cleanroom using a triple-wrap system. The outer layer is removed in the material airlock, the second layer in the gowning room, and the final layer only at the point of use (refer to our dedicated chapter below on components and suppliers). Dedicated waste removal: waste should be collected in designated, cleanroom-compatible containers and removed following a specific protocol that prevents it from contaminating clean areas on its way out. |
9. Personal ItemsAnything not essential to the process is a source of contamination. Zero tolerance for personal effects: this includes jewelry (even under gloves), watches, cell phones, keys, and any paper or writing instruments that are not specifically designed for cleanroom use. Approved cleanroom supplies: only use cleanroom-certified paper, pens, and tools that are designed to be non-shedding. |  |
10. Health and Wellness PoliciesAn operator’s health directly affects the cleanroom environment. Stay home when sick: policies must be in place that encourage personnel to report illnesses like colds, coughs, or skin conditions and stay home. Coming to work sick poses a massive contamination risk.  Cover all skin lesions: any cuts, abrasions, or sunburns must be completely covered with a non-shedding, approved bandage before gowning. | 11. Gloved Hands RoutineGloves are the primary point of contact with equipment and materials and can easily become cross-contaminated. Sanitize before and after key tasks: gloved hands should be sprayed with a sterile 70% IPA solution and allowed to air dry before handling critical products, after touching non-critical surfaces (like equipment panels), and at regular intervals. Proper sanitization technique: ensure the entire surface of both gloves is thoroughly wetted with the sanitizing agent. The industry secret (that nobody is proud about): gloves, used in single or even double layers are hermetic … making operators sweating. And because they are single use and thin … they are fragile. So where does the sweat go once the glove brakes? Have a strict glove-change policy, so as the room temperature control to limit sweating. |
 | 12. Gowning Room as a Transition ZoneThe gowning room is not a break room; it is a critical control point. No socializing: the gowning room is for the sole purpose of gowning and de-gowning. Social conversations should not take place there. No coffee machine here! Follow the demarcation line: a “sit-over bench” or similar demarcation separates the “dirty” side from the “clean” side. Personnel must never allow their “dirty” side footwear or garments to touch the “clean” side floor. |
Tip: as likely only the eyes should be visible for a correctly dressed operator, it can be necessary to have their individual names on the clothing -but without velcro- (to be balanced with the fact that they should be as little operators as possible, and they should know each other well).
Personnel Conduct, Qualification, Personal Accountability and Procedural Adherence
And more generally, as typical in those regulated industries for any process: all personnel must be formally qualified prior to entry into any classified area. Qualification is achieved through a documented training program covering area-specific SOPs, aseptic behaviors, and the principles of contamination control. This training shall instill a state of constant situational awareness, ensuring personnel understand the specific requirements of their operational zone.
Suppliers Specifications
Will only be detailed here some typical examples of specific instructions to suppliers intended to delivers goods for cleanroom usage.
Tip: be ready for a steep learning curve for your suppliers if they are not already used to deliver for cleanrooms (own experience: just start by explaining that you need a special material for the labels and that just “sticking” is not enough …).
Note: “usual” regulated industries specifications are not included here (supplied materials must come with a Certificate of Conformance (CoC) and a Certificate of Analysis (CoA) that specifies the material composition, lot number, and compliance with relevant standards. Full traceability from raw material to finished component is required, with records maintained and available for audit).
1. Mandated Material Composition and Surface Finish
All components must be fabricated from materials that are inherently low-particulating and low-outgassing.
- Forbidden materials: absolutely no raw wood, paper-based products, uncoated carbon steel, natural-fiber textiles (e.g., cotton, wool), or materials containing powders or silicones are permitted in the final component.
- Surface requirements: surfaces must be non-porous and smooth to prevent particle shedding and to facilitate cleaning. For metals, a finish such as electropolishing is required. For plastics, a molded finish free of flash and burrs is necessary. Specify a maximum surface roughness (e.g., Ra < 0.8 µm) where applicable.
2. Validated Precision Cleaning Process
All goods must undergo a final, validated cleaning process immediately before packaging.
- Methodology: this typically involves a multi-stage ultrasonic cleaning process using detergents approved for cleanroom use, followed by multiple cascade rinses with heated, high-purity Deionized (DI) water.
- Final rinse verification: the final DI water rinse must be tested for quality. You must provide data showing the final rinse water meets specifications for resistivity (e.g., >18 MΩ·cm), Total Organic Carbon (TOC), and Liquid Particle Count (LPC) to prove all residues and foreign particles have been removed.
- Drying protocol: parts must be dried using a non-contaminating method, such as blowing with heated, HEPA- or ULPA-filtered nitrogen or clean dry air. Wiping with any type of cloth, even “low-lint” wipes, is forbidden at this final stage.
3. The Double-Bagging Packaging Mandate
All items must be double-bagged inside a certified cleanroom environment. This protocol is non-negotiable and must be followed precisely.

- Packaging environment: the inner bag must be sealed within an environment certified to be as clean or cleaner than the end-user’s cleanroom (e.g., an ISO 5 environment for goods going into an ISO 5 cleanroom).
- Bag material: only use new, certified cleanroom-grade polyethylene or nylon bags. The bags must be low in static propensity and free from processing additives like slip agents or plasticizers. Provide certification for the bags themselves.
- Inner bag sealing: place the cleaned, dried parts into the inner bag. If specified, purge the bag with filtered nitrogen to provide a cushioned, inert atmosphere. Seal the bag using a thermal impulse sealer to create a continuous, robust seal. No tape, staples, or zip-closures are permitted.
- Inter-stage wipe down: before passing to the next packaging stage, the exterior of the sealed inner bag must be meticulously wiped down with a 70/30 solution of Isopropyl Alcohol (IPA) and DI water using a sterile, non-shedding polyester wipe.
- Outer bag sealing: in an environment of a lower classification (e.g., ISO 7), place the wiped-down inner bag into the outer bag and seal it in the same manner. This creates a system where the outer bag can be removed in the gowning room or material airlock, allowing a clean inner bag to enter the main cleanroom.
Expert tip: in some cases, if the inner bag is air-tight sealed, mind the pressure inflating effect if it needs to be air shipped (sealing or cardbox around strong enough, or leave room for expansion …)
4. Strict “No-Paper, No-Cardboard” Rule for Packaging

No paper or cardboard can be in direct contact with the component or its primary/secondary cleanroom bags.
- Labels: all identification labels applied directly to the bags must be made of synthetic material (e.g., polyester) with an acrylic adhesive and printed with non-shedding ink. The label must be placed on the exterior of the outer bag. If an internal “traveler” tag is required, it must be printed on cleanroom-grade synthetic paper (e.g., Tyvek).
- Documentation: all paperwork, such as Certificates of Conformance, analysis reports, and packing slips, must be placed in a separate pouch attached to the exterior of the final, sealed shipping box or sent electronically. Under no circumstances should any paper be placed inside the shipping box itself.
5. Final Shipping Container Preparation
The final shipping container (typically a cardboard box) is a source of gross contamination and must be prepared to isolate the clean-packaged goods.
- Box liner: the shipping box must first be lined with a fresh, general-purpose polyethylene liner bag.
- Dunnage: use only cleanroom-compatible dunnage, such as new polyethylene foam or bubble wrap, to cushion the double-bagged product inside the box liner. The use of packing peanuts, shredded paper, or kraft paper is strictly forbidden.
- Sealing: after placing the double-bagged product inside the lined box, fold the liner over and then seal the cardboard box with standard packing tape. This ensures that when the box is opened, the first thing the receiver touches is the clean liner, not cardboard dust.
Outro

A robust Contamination Control Strategy is not merely a document, but the active intelligence governing all cleanroom operations. It translates quality risk management principles into the tangible, daily execution of best practices, from material transfer and aseptic technique to validated disinfection protocols.
Ultimately, the success of the strategy is measured not by its scope, but by its verifiable outcomes—the consistent data from environmental monitoring that confirms a state of control and informs the continuous evolution of the system.
FAQ
How does a Contamination Control Strategy (CCS) differ from individual cleaning and gowning SOPs?
A CCS is the overarching strategic document that integrates all control elements based on risk. SOPs are the detailed, tactical instructions for executing specific tasks defined within that strategy.
What is the correct technique for wiping items for transfer into a Grade A/B area, and why is contact time critical?
The correct technique involves using sterile, low-lint wipes saturated with a validated disinfectant in overlapping, unidirectional strokes. Contact time is critical because it is the validated duration required for the disinfectant to achieve its sporicidal or bactericidal action.
What are the most critical data inputs for the periodic review and update of a CCS?
The most critical inputs are environmental and personnel monitoring trend data, process deviations, media fill results, and CAPA effectiveness data. These inputs verify control and highlight areas needing reassessment.
What is the best practice for integrating supplier and raw material controls into the facility’s CCS?
The CCS must reference the supplier qualification program, which mandates audits, quality agreements, and microbial specifications for incoming materials. This extends contamination control to the entire supply chain.
Why are unidirectional, overlapping strokes mandated for cleaning and disinfecting surfaces in a cleanroom?
This technique ensures complete surface coverage and prevents the re-contamination of areas that have already been cleaned. It physically lifts and removes contaminants rather than just spreading them around the surface.
What are the key validation requirements for cleaning and disinfection agents cited in a CCS?
Validation requires efficacy studies (coupon studies) using in-house microbial isolates on representative manufacturing surfaces. These studies must prove the effectiveness of the agents at their specified contact times.
What are the primary differences in a CCS for an aseptic process versus a terminally sterilized product?
An aseptic process CCS focuses intensely on the exclusion of all microorganisms, as there is no final kill step. A CCS for terminal sterilization focuses on controlling the pre-sterilization bioburden to a validated limit.
What is the correct sequence for de-gowning, and why is it as critical as the gowning process itself?
De-gowning proceeds from the “dirtiest” items to the “cleanest,” typically starting with gloves and moving inward to the coverall, to contain contaminants on the gown. This prevents the operator from contaminating their scrubs or skin and dispersing particles from the used gown into the changing room.
Related Readings
- Aseptic Process Simulation (Media Fill) design and interpretation: it involves designing worst-case simulations to qualify a process’s aseptic integrity. Interpreting the results is critical for validating the process and qualifying operators.
- Microbial identification strategies and data trending: this deals with identifying environmental isolates to the species level to understand the facility’s microflora. Trending this data is essential for detecting shifts that indicate a loss of control.
- Design and control of RABS and isolator technology: it explores the validation and operation of advanced barrier systems. It includes integrity testing and glove management to ensure a superior aseptic environment.
- WFI and Pure Steam System validation and biofilm control: it shall cover the design and routine monitoring of high-purity water systems. A primary focus is on system validation and ongoing strategies to prevent and control biofilm formation.
- Container Closure Integrity Testing (CCIT) methodologies: the technologies used to ensure the final product seal is integral, guaranteeing sterility until use. It includes methods like vacuum decay and high voltage leak detection.
- Supplier and raw material microbial control programs: it shall extend contamination control to the supply chain through supplier audits and quality agreements. It involves setting and verifying microbial specifications for all incoming materials.
- Sterilization validation for components and equipment: validating the lethality and repeatability of sterilization cycles (e.g., autoclave, dry heat) for all items entering the aseptic processing area.
- Management of disinfectant and cleaning agent residues: it addresses the potential for chemical residues to inhibit disinfectant efficacy or become a product contaminant. It covers strategies for residue detection, removal, and rotation of cleaning agents.
External Links on Contamination Control Strategy (CCS)
International Standards
- ISO 14644-1: 2015 Cleanrooms and controlled environments – Part 1: Classification of air cleanliness
- ISO 14644-2: 2015 Cleanrooms and controlled environments – Part 2: Monitoring to provide evidence of cleanroom performance related to air cleanliness by airborne particulate contamination
- ISO 14644-3: 2019 Cleanrooms and controlled environments – Part 3: Test methods
(hover the link to see our description of the content)
Glossary of Terms Used
Bioburden: the presence of viable microorganisms on a surface or in a substance, typically measured to assess contamination levels in pharmaceuticals, medical devices, and other sterile products. It is crucial for determining sterilization efficacy and product safety.
Building Management System (BMS): a centralized control system that monitors and manages a building's mechanical and electrical equipment, including heating, ventilation, air conditioning, lighting, security, and fire safety systems, to enhance operational efficiency, comfort, and energy management.
Carbon Capture & Sequestration (CCS): a process that captures carbon dioxide emissions from sources like power plants and industrial processes, transporting it for storage underground or utilizing it in various applications, thereby reducing greenhouse gas concentrations in the atmosphere.
Certificate of Analysis (CoA): a document issued by a manufacturer or testing laboratory that confirms a product's specifications, quality, and compliance with regulatory standards, detailing test results and methods used for analysis.
Certificate of Conformance (CoC): a document issued by a manufacturer or supplier confirming that a product meets specified standards, regulations, or contractual requirements, often used in quality assurance and compliance verification processes.
Code of Federal Regulations (CFR): a compilation of the general and permanent rules published by federal agencies in the United States, organized by subject matter into 50 titles, serving as the official legal source for federal regulations.
Colony Forming Units (CFU): a measurement used to estimate the number of viable microorganisms in a sample, indicating the number of cells capable of forming colonies under specific growth conditions.
Computational Fluid Dynamics (CFD): a numerical method used to analyze fluid flow, heat transfer, and related phenomena by solving the governing equations of fluid motion and thermodynamics through discretization techniques, enabling simulation and visualization of complex fluid behavior in various engineering applications.
Contamination Control Strategy (CCS): a systematic approach to prevent, detect, and mitigate contamination in controlled environments, ensuring product quality and safety through defined procedures, monitoring, and risk management practices.
Corrective Action and Preventative Action (CAPA): a systematic approach to identifying, investigating, and addressing nonconformities and potential issues to prevent recurrence and ensure compliance with regulatory standards in quality management systems.
Corrective and Preventive Action (CAPA): a systematic approach to identifying, investigating, and addressing nonconformities to prevent recurrence and eliminate potential issues, ensuring compliance with regulatory standards and continuous improvement in processes and products.
Critical Control Points (CCP): specific stages in a process where control can be applied to prevent, eliminate, or reduce food safety hazards to acceptable levels. Identifying these points is essential for effective hazard analysis and critical control management in food production systems.
current Good Manufacturing Practice (cGMP): a system ensuring that products are consistently produced and controlled according to quality standards, encompassing regulations and guidelines for manufacturing processes, facilities, equipment, and personnel to ensure safety, quality, and efficacy in pharmaceuticals, food, and other regulated industries.
Electronics Manufacturing Services (EMS): outsourced services that provide design, production, assembly, and testing of electronic components and systems for original equipment manufacturers, enabling cost-effective and efficient manufacturing processes while allowing clients to focus on core business activities.
Environmental Monitoring System (EMS): a system designed to systematically collect, analyze, and report data on environmental conditions, including air, water, and soil quality, to assess compliance with regulations, track changes over time, and support decision-making for environmental management and protection.
Failure Mode and Effects Analysis (FMEA): a systematic method for evaluating potential failure modes within a system, process, or product, assessing their effects on performance, and prioritizing risks to improve reliability and safety through corrective actions.
Food and Drug Administration (FDA): a federal agency of the United States Department of Health and Human Services responsible for regulating food safety, pharmaceuticals, medical devices, cosmetics, and tobacco products to ensure public health and safety through scientific evaluation and enforcement of compliance standards.
Good Manufacturing Practice (GMP): a system ensuring products are consistently produced and controlled according to quality standards, minimizing risks involved in pharmaceutical production and related industries. It encompasses guidelines for manufacturing processes, facility conditions, personnel qualifications, and documentation practices to ensure product safety and efficacy.
Hazard Analysis and Critical Control Points (HACCP): a systematic approach to food safety that identifies, evaluates, and controls hazards at critical points in the production process to prevent foodborne illnesses and ensure product safety.
Heating Ventilation and Air Conditioning (HVAC): a system designed to regulate indoor climate by controlling temperature, humidity, and air quality through heating, cooling, and ventilation processes. It includes components such as furnaces, air conditioners, ductwork, and thermostats for efficient environmental management.
Heating, Ventilation, and Air Conditioning (HVAC): a system designed to regulate indoor climate by controlling temperature, humidity, and air quality through heating, cooling, and ventilation processes, ensuring comfort and safety in residential, commercial, and industrial environments.
International Organization for Standardization (ISO): a non-governmental international body that develops and publishes standards to ensure quality, safety, efficiency, and interoperability across various industries and sectors, facilitating global trade and cooperation. Established in 1947, it comprises national standardization organizations from member countries.
Key Performance Indicator (KPI): a measurable value that demonstrates how effectively an organization is achieving key business objectives, often used to evaluate success at reaching targets.
Material Airlock (MAL): a sealed entryway designed to control the transfer of materials between different environments, preventing contamination and maintaining specific atmospheric conditions. It typically consists of two or more interlocking doors that ensure isolation during the transfer process.
Performance Qualification (PQ): a process that verifies a system or equipment operates according to specified requirements under real-world conditions, ensuring it consistently performs its intended function within predetermined limits.
Personnel Airlock (PAL): a sealed entryway designed to allow personnel to transition between different pressure environments while minimizing contamination and maintaining safety, typically used in space stations, laboratories, or cleanrooms. It features interlocking doors that prevent simultaneous opening.
Standard Operating Procedure (SOP): a set of step-by-step instructions created to help workers carry out routine operations consistently and efficiently, ensuring compliance with regulations and quality standards.
Statistical Process Control (SPC): a method of quality control that employs statistical techniques to monitor and control a process, ensuring it operates at its full potential by identifying variations and maintaining consistent output within specified limits.
Total Organic Carbon (TOC): a measure of the amount of carbon found in organic compounds within a sample, often used to assess water quality and environmental health. It quantifies the concentration of carbon derived from organic matter, excluding inorganic carbon sources.
Unique Selling Point (USP): a distinctive feature or benefit that sets a product or service apart from competitors, aimed at attracting customers by addressing specific needs or preferences.
Volatile Organic Compound (VOC): organic chemicals that have a high vapor pressure at room temperature, leading to significant evaporation and potential air pollution. They are commonly found in paints, solvents, and fuels, contributing to smog formation and adverse health effects.